In the landscape of neurodegenerative medicine, Huntington’s disease has long stood as a formidable, multifaceted puzzle. Characterized by involuntary movements, cognitive decline, and psychiatric disturbances, this fatal, inherited condition has been the subject of intense scrutiny for decades. Now, a groundbreaking study published in the journal Cell in 2026 has provided a vital missing piece to the puzzle, revealing that a singular, slow-moving genetic mechanism may be the culprit behind damage across multiple, distinct regions of the brain.
Researchers from the Broad Institute of MIT and Harvard, Harvard Medical School, and McLean Hospital have identified that the toxic expansion of DNA repeats—the hallmark of the disease—operates in the neocortex just as it does in the striatum. This finding suggests that emerging therapeutic strategies targeting this genetic instability could provide a comprehensive shield for the brain, potentially addressing both motor and cognitive decline simultaneously.
The Main Facts: A Unified Mechanism of Toxicity
For years, scientists have understood that Huntington’s disease is triggered by a genetic stutter: an inherited expansion of a three-letter DNA sequence, "CAG," within the Huntingtin (HTT) gene. While healthy individuals typically carry fewer than 36 of these repeats, those who inherit 40 or more are at high risk of developing the disease.
However, the "why" and "how" of the transition from a genetic predisposition to clinical manifestation remained unclear. In 2025, a landmark study from the Broad Institute revealed that the HTT gene does not remain static. Instead, it slowly expands over a person’s lifetime, accumulating repeats until it crosses a catastrophic threshold—roughly 150 repeats—at which point the gene becomes toxic, causing cellular dysfunction and death.
Initially, this "threshold theory" was observed only in the striatum, the region of the brain that governs movement and is the first to succumb to the disease. The new 2026 study confirms that this same biological "ticking time bomb" is present in the neocortex, the region responsible for higher-level cognitive functions. By analyzing over 125,000 individual cells from human brain tissue, the researchers demonstrated that cortical cells follow an identical path to toxicity, even if they arrive there at a different pace.
Chronology of Discovery: From Observation to Insight
The trajectory of this research represents a shift from viewing Huntington’s as a collection of localized brain failures to understanding it as a systemic, gene-driven process.
The Decades-Long Consensus
Since the HTT gene was first identified in the 1990s, the field focused largely on the presence of the expanded gene itself. Researchers debated whether the protein produced by the gene was inherently toxic from birth or if something changed over time. The consensus was that the mutation was permanent and static.
The 2025 Paradigm Shift
The 2025 study challenged this long-held dogma by showing that the HTT gene is dynamic. Using advanced genomic sequencing, researchers observed that the DNA repeats expand in somatic cells throughout the patient’s life. The realization that the mutation grows—and that it only becomes toxic once it reaches a critical mass of ~150 repeats—upended the field’s understanding of the disease’s progression.
The 2026 Expansion
The current research sought to determine if this mechanism was unique to the striatum or a broader, brain-wide phenomenon. By applying the same high-resolution single-cell analysis to the neocortex, the team confirmed that the 150-repeat threshold is a universal trigger for toxicity. Furthermore, they identified that while the trigger is the same, the tempo of the expansion differs, which helps explain why some areas of the brain show symptoms earlier than others.
Supporting Data: Peering Inside the Neuron
To reach these conclusions, the research team conducted a massive, high-resolution audit of human brain tissue. They analyzed 125,000 cells harvested from the neocortex of eleven individuals who had lived with Huntington’s disease.
The Threshold Effect
The data revealed a striking consistency: in both the striatum and the cortex, gene expression—the process by which DNA instructions are converted into functional proteins—began to fail only after the HTT gene crossed the 150-repeat threshold. Before this threshold, cells appeared relatively stable; after it, they descended into a state of molecular chaos.
The Pace of Progression
A critical nuance discovered in the 2026 study is the variable rate of expansion. In the neocortex, the expansion process is significantly slower than in the striatum. Many cortical cells, in fact, never reach the toxic 150-repeat threshold during the person’s lifespan, which explains why certain neural pathways remain functional longer than others. This "slow-burn" hypothesis provides a compelling explanation for the heterogeneous nature of Huntington’s symptoms.
The Mystery of Nuclear Inclusions
For decades, researchers have debated the role of "nuclear inclusions"—clumps of mutant protein found inside the nuclei of brain cells. Some believed they were the primary cause of damage, while others argued they were protective, acting as a "trash bin" for harmful proteins.
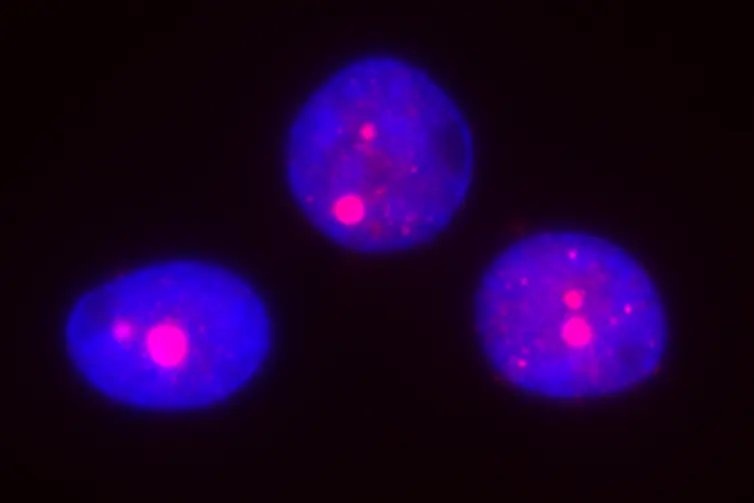
The new study provides a definitive answer: nuclear inclusions are a final, desperate indicator of cellular distress. They were only found in cells that had already surpassed the 150-repeat threshold and were nearing death. These inclusions are essentially "red flags" signaling that a neuron is in the final stages of dysfunction.
Official Responses: A Call for Urgency
The implications of these findings have resonated throughout the scientific community, providing a sense of renewed hope for therapeutic intervention.
Steve McCarroll, a co-senior author of the study and a prominent geneticist at the Broad Institute, emphasized the clinical potential of these findings. "These new results suggest that such medicines, if and when they’re successfully developed, might be able to address not just one of the pathologies in the disease, but the two big ones," McCarroll stated. By targeting the expansion process itself, scientists may be able to prevent cells from ever reaching the toxic threshold, effectively halting the disease at its source.
Sabina Berretta, co-senior author and director of the Harvard Brain Tissue Resource Center at McLean Hospital, underscored the importance of this discovery for patient care. "By pointing to a shared disease mechanism across brain regions, our findings suggest these new efforts are on the right track," Berretta noted. "We hope this evidence adds urgency to developing such treatments."
The researchers were also careful to acknowledge the human cost of the disease and the profound contributions of those who donated their brain tissue to science. "Above all, this work is a testament to the extraordinary generosity, commitment, and foresight of brain donors and their families, whose contributions made these discoveries possible," the team collectively remarked.
Implications: The Future of Huntington’s Therapeutics
The identification of a shared mechanism across the striatum and the neocortex is a "game-changer" for drug development.
Broad-Spectrum Treatments
If a medicine is designed to stabilize the HTT gene or prevent the expansion of CAG repeats, it no longer needs to be localized to one part of the brain. The fact that the same mechanism is active in multiple regions suggests that a single systemic treatment could theoretically protect the entire brain, addressing the full spectrum of motor, cognitive, and psychiatric symptoms.
Precision Medicine and Biomarkers
The identification of nuclear inclusions as a sign of terminal cellular dysfunction provides clinicians with a potential biomarker. If doctors can monitor the accumulation of these inclusions or the expansion of repeat lengths in patient cells, they may be able to better track disease progression and measure the efficacy of new experimental drugs.
Shifting the Focus to Prevention
Perhaps the most significant implication is the shift toward preventative medicine. If scientists can find a way to intervene before a cell crosses the 150-repeat threshold, they may be able to extend the lives and quality of life for those at risk of Huntington’s. By slowing or stopping the "genetic clock," the disease could potentially be transformed from a rapidly progressing, fatal condition into a manageable one.
As the scientific community moves forward, the focus will now shift to translating these molecular findings into pharmacological solutions. While the journey from the lab bench to the clinic is often long, the clarity provided by this study—that Huntington’s is governed by a predictable, universal genetic trigger—provides the most promising roadmap yet for those dedicated to curing this devastating disease.
About the Author