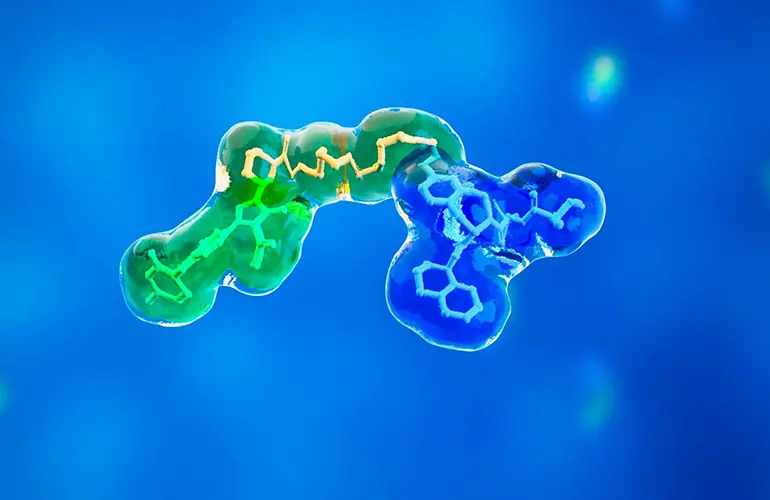
The landscape of modern pharmacology was fundamentally altered in May 2026 with the U.S. Food and Drug Administration’s (FDA) approval of vepdegestrant (marketed as Veppanu®). As the first-ever Proteolysis Targeting Chimera (PROTAC) to receive regulatory clearance, its arrival serves as a watershed moment for the pharmaceutical industry. This milestone validates the "chemical knockdown" modality, a sophisticated approach that targets proteins previously deemed "undruggable" by traditional occupancy-based inhibitors. However, the path from benchtop discovery to the pharmacy shelf for PROTACs remains fraught with significant developmental hurdles—most notably, the challenge of achieving effective oral bioavailability.
Main Facts: The Mechanism of Chemical Knockdown
PROTACs represent a paradigm shift in how we treat disease. Unlike traditional small-molecule drugs that function by occupying a protein’s active site to inhibit its function, PROTACs act as catalytic "matchmakers." A typical PROTAC molecule is a heterobifunctional construct comprising three critical components:
- A target-binding warhead: A moiety that identifies and binds to the disease-causing protein.
- An E3 ligase recruiter: A component that binds to an E3 ubiquitin ligase.
- A chemical linker: A flexible tether that connects the two ends, allowing the molecule to bring the target protein into proximity with the E3 ligase.
Once this ternary complex is formed, the E3 ligase tags the target protein with ubiquitin molecules, marking it for destruction by the cell’s internal disposal system, the ubiquitin-proteasome system (UPS). This "event-driven" mechanism is catalytic; once the target is degraded, the PROTAC is released to repeat the cycle. Because the drug does not need to occupy 100% of the target protein to be effective, it can maintain therapeutic efficacy at much lower concentrations than traditional inhibitors, potentially overcoming the resistance mechanisms that often plague conventional therapies.
Chronology: From Concept to Clinical Reality
The evolution of PROTAC technology has been a multi-decade journey of innovation in medicinal chemistry:
- Early 2000s: The foundational concept of "Targeted Protein Degradation" (TPD) was pioneered, demonstrating that E3 ligases could be hijacked to degrade specific proteins.
- 2010s: The field saw a rapid expansion as researchers optimized bifunctional degraders, moving from proof-of-concept experiments in cell cultures to robust in vivo efficacy studies.
- Early 2020s: Heavy investment poured into TPD platforms, focusing on optimizing the "beyond-Rule-of-Five" (bRo5) chemical space that these large molecules occupy.
- May 2026: The FDA grants approval for vepdegestrant, confirming that the logistical and pharmacological challenges of this class are surmountable.
Despite this success, the clinical pipeline remains characterized by a high attrition rate for orally administered candidates. Developers are now shifting their focus toward the "early-phase" strategy—identifying and mitigating oral exposure limitations long before a candidate reaches a pivotal trial.
Supporting Data: The "Beyond Rule of Five" Dilemma
The primary hurdle for oral PROTAC development is rooted in the "Rule of Five" (Ro5), the heuristic used to predict the oral bioavailability of small molecules. Ro5 suggests that compounds with a molecular weight >500 Da, high hydrogen-bond donor (HBD) counts, and high lipophilicity are less likely to be orally active.
PROTACs almost invariably violate these rules. They are frequently characterized by:
- Molecular weights exceeding 700–800 Da.
- High polar surface area (PSA > 150 Ų).
- An excessive number of rotatable bonds (>15).
These attributes lead to poor membrane permeability and solubility. Research published in 2024 suggests that simply applying Ro5 is insufficient. Instead, developers must look at "solvent-exposed hydrogen-bond donors" (eHBD). Data indicates that limiting eHBD to ≤2 can be a powerful discriminator in selecting candidates with better oral potential. Furthermore, because these molecules often fail due to a combination of poor solubility and limited permeability, they require a cross-functional approach that integrates DMPK (drug metabolism and pharmacokinetics), structural biology, and advanced bioanalysis.
Strategies to Solve the Oral Exposure Problem
To optimize a PROTAC for oral delivery, developers are employing several sophisticated strategies, each designed to balance the molecule’s biological activity with its pharmacokinetic profile.
1. Decoding the Food Effect
Many PROTACs exhibit solubility limitations that can be mitigated by the patient’s dietary state. Biorelevant media studies (using FaSSIF/FeSSIF) have shown that certain degraders exhibit significantly improved solubility in a fed-state environment. Vepdegestrant’s clinical label, which specifies dosing with food, is a testament to this strategy. Developers should not view the food effect as a limitation, but as a parameter to be characterized early in the ADME (absorption, distribution, metabolism, and excretion) profiling process.
2. Strategic Linker Engineering
The linker is the "engine room" of the PROTAC. It dictates not only the orientation required for degradation but also the molecule’s passive permeability. By replacing traditional PEG linkers with more rigid structures—such as 1,4-disubstituted phenyl rings—or by reducing the number of amide bonds, researchers can improve the molecule’s ability to cross intestinal membranes. The current gold standard is to design linkers that encourage the molecule to adopt a "folded" or "collapsed" conformation in non-polar environments, which shields polar groups and enhances membrane diffusion.
3. E3 Ligase Selection
The choice of E3 ligase is not merely a biological decision; it is a developability decision. While there are many E3 ligases, CRBN (Cereblon) and VHL remain the industry favorites. CRBN-based degraders often allow for smaller, more "drug-like" molecular footprints compared to those recruiting larger E3 complexes. This makes CRBN an attractive scaffold for teams prioritizing oral bioavailability from the onset of lead optimization.
4. The Power of Intramolecular Hydrogen Bonding (IMHB)
By engineering the molecule to form internal hydrogen bonds, scientists can effectively "hide" polar features from the solvent, reducing the effective polar surface area. This folding mechanism is essential for navigating the complex journey through the gut lining and into the target cell. Computational modeling and structural biology are now playing an outsized role in predicting which molecular architectures will favor this folded, membrane-permeable state.
5. Prodrugs and Molecular Glues
When structural modifications to the PROTAC fail to yield sufficient exposure, developers are turning to prodrug strategies. By masking polar groups with lipophilic "promoieties" that are cleaved only after absorption, researchers can bypass early metabolic or solubility hurdles. Alternatively, if a PROTAC remains too bulky or unstable, the field is increasingly exploring "molecular glues." These agents are smaller and simpler than PROTACs; they do not use a linker but instead induce a change in the protein surface that promotes degradation. While not a replacement for PROTACs, they offer a viable alternative for targets where the bifunctional architecture proves too burdensome for oral delivery.
Implications for the Future of Drug Development
The approval of vepdegestrant has shifted the goalposts for the entire pharmaceutical industry. The focus has moved from "Can we degrade this protein?" to "Can we deliver this degradation modality effectively as an oral pill?"
For drug sponsors, the implications are clear: the "siloed" approach to drug development is dead. Successfully advancing an oral PROTAC requires the integration of medicinal chemistry with early-stage DMPK and bioanalytical studies. If a team waits until late-stage preclinical development to address solubility or permeability, the cost of failure becomes prohibitive.
Furthermore, the rise of TPD highlights the growing importance of "fit-for-purpose" bioanalysis. Because PROTACs have unique pharmacokinetic profiles—where the drug concentration in the plasma may not perfectly correlate with the duration of the target degradation—developers must invest in ultrasensitive assays that measure not just drug concentration, but the downstream pharmacodynamic effect (the "knockdown" itself).
As we look toward the remainder of the decade, the industry is poised to see a surge in oral PROTAC programs. The lessons learned from early pioneers—that linker rigidity, E3 ligase selection, and the strategic use of IMHB are the keys to the kingdom—will provide the blueprint for the next generation of targeted therapies. While the chemical space is "beyond the rule of five," the path forward is becoming increasingly well-defined, promising a future where the most difficult-to-treat diseases are addressed by the most precise technology in the history of medicine.
Tao Xiong, Director in the DMPK Department at WuXi AppTec, emphasizes that the key to success lies in early identification of liabilities. With nearly two decades of experience, Xiong notes that the industry’s ability to support over 200 IND-related programs in this space is a testament to the robust, interdisciplinary workflows now standard in modern drug discovery. The transition from intravenous, specialized administration to patient-friendly oral dosing is the final frontier in making PROTACs the standard of care for oncology and beyond.
About the Author