The landscape of modern pharmacology was irrevocably altered by the arrival of AlphaFold. By solving the long-standing "protein folding problem," the tool provided a structural blueprint for the building blocks of life, effectively opening a door that had been locked for decades. However, as the initial euphoria surrounding predictive structural biology wanes, a sobering reality has set in: knowing the shape of a protein is not the same as knowing how to disable it with a drug.
Seattle-based biotechnology firm Talus Bio is now challenging the hegemony of structure-based drug design. By abandoning the pursuit of three-dimensional protein maps in favor of a novel, mass-spectrometry-informed approach, the company claims to have achieved a 5,000-fold increase in screening speed. Their new platform, Ptarmigan-1, is targeting the "undruggable" class of proteins—specifically transcription factors—that have long frustrated oncologists and medicinal chemists alike.
The Structural Bottleneck: Why AlphaFold Isn’t Enough
To understand the magnitude of Talus Bio’s pivot, one must first recognize the limitations of current paradigms. While AlphaFold and its successors have mapped hundreds of millions of proteins, these models excel primarily with rigid, globular structures. They function on the assumption that a protein has a "lock" into which a drug "key" can be inserted.
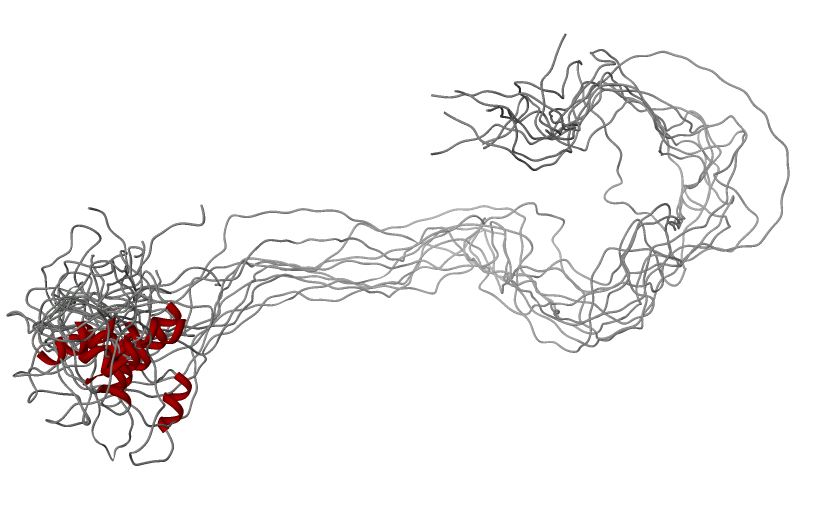
However, a vast swath of the human proteome—roughly 50%, according to Talus CTO Lindsay Pino—does not conform to this rigid geometry. Many proteins, particularly transcription factors which govern the activation of genes, are characterized by "intrinsically disordered regions." These proteins exist in a state of constant, fluid motion.
"If you try to fold a transcription factor, you just get a plate of spaghetti," says Pino. "You can’t do drug discovery on a plate of spaghetti."
Because these proteins lack a fixed, stable pocket, traditional structure-based virtual screening fails. This failure is a primary contributor to the fact that nearly 87% of the 20,000 known human proteins remain untouched by approved small-molecule drugs. The industry has effectively been ignoring half of the potential therapeutic targets in the human body simply because current software cannot "see" them.
A Chronology of the Shift: From Geometry to Data-Driven Engagement
The development of Ptarmigan-1 represents a strategic departure from the industry’s obsession with 3D visualization.
The Era of Geometric Prediction
For years, the gold standard for drug discovery involved identifying a potential target, using cryo-electron microscopy or X-ray crystallography to determine its structure, and then using computational docking to see which drug candidates might fit into that structure. This process is time-consuming, expensive, and often impossible for flexible targets.
The Rise of the "Structure-Free" Approach
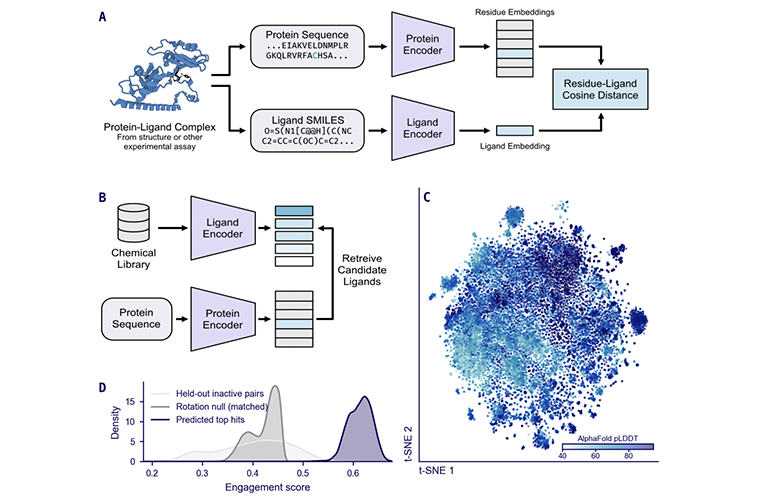
Talus Bio, under the leadership of CEO Alex Federation and CTO Lindsay Pino, began investigating an alternative: what if they didn’t need the shape to predict the binding? Their research, formalized in a July 2026 preprint, posits that mass spectrometry data—which records the actual physical engagement between compounds and proteins inside a living cell—contains more useful information than a theoretical 3D model.
The Benchmark and Validation
In recent months, the team tested the efficacy of Ptarmigan-1 against established benchmarks. In a retrospective study on STAT6, a notoriously difficult transcription factor involved in various inflammatory conditions and cancers, the AI was tasked with identifying inhibitors from a library of patents. While traditional docking models and contemporary AI tools like Boltz-2 performed no better than chance (scoring 0.58), Ptarmigan-1 achieved an area under the curve (AUC) of 0.94, successfully identifying 40 known inhibitors.
Supporting Data: The 5,000-Fold Advantage
The most compelling argument for the Talus Bio approach is not just accuracy, but computational efficiency. In the world of high-throughput screening, time is synonymous with capital.
The Cost of Complexity
Federation highlights the staggering economic burden of modern virtual screening. "A big experiment in drug discovery in the real world is a million compounds," he explains. "That would cost something like $100,000 to $1 million and take months, for one protein. Folding the protein is the computational expense."
Efficiency Metrics
When comparing the performance of Ptarmigan-1 against Boltz-2 on a single Nvidia H100 GPU, the results were transformative:
- Boltz-2: Required approximately 54 seconds per compound.
- Ptarmigan-1: Required only 10 milliseconds per compound.
Extrapolating these figures, a screen of one million compounds that would have taken nearly two years using structural prediction methods can now be completed by Ptarmigan-1 in under three hours. In one demonstration, the team searched a library of 3.4 billion compounds against all 20,431 human proteins in under 24 hours, utilizing only 20 H100 GPU-hours.
Official Perspectives: The Synergy of Approaches
Despite the competitive nature of the field, the leadership at Talus Bio does not advocate for the total abandonment of structural models. Instead, they propose a complementary ecosystem.
"We’ve been sitting on these targets we’ve known about for decades, that we know are good targets," says Pino. "We just haven’t had the tools to find them yet."
The company suggests that the ideal workflow involves using Ptarmigan-1 as a "funnel." By rapidly screening billions of compounds to narrow the field down to the most promising candidates, researchers can then apply more intensive, structure-based models to the survivors. This hybrid strategy allows for the speed of data-driven screening with the precision of structural refinement.
The implications for the industry are profound. By removing the need for structure, Talus Bio has essentially democratized the ability to screen "undruggable" proteins. This could trigger a massive influx of new drug candidates into the development pipeline, particularly for rare diseases and complex cancers where transcription factors play a central role.
Implications: The Future of Small-Molecule Design
The success of Ptarmigan-1 signals a potential shift in the "AI for drug discovery" narrative. For the past decade, the industry has chased the dream of "the perfect model"—one that could simulate every atomic interaction with perfect fidelity. Talus Bio’s findings suggest that, in many cases, we may have been over-engineering the problem.
1. Accelerating Time-to-Clinic
If the time required to identify a lead molecule drops from months to hours, the bottlenecks in drug discovery shift from discovery to clinical testing. This could allow for more agile experimentation, where researchers can pivot between targets as new biological insights emerge, rather than being tethered to a single, multi-year project.
2. Redefining the "Undruggable"
The term "undruggable" has always been a limitation of our tools, not necessarily the biology. By proving that binding can be identified without a 3D pocket, Talus is forcing a re-evaluation of the entire human proteome. Targets that were shelved because they were too "floppy" or "disordered" are suddenly viable candidates for high-throughput screening.
3. Computational Democratization
The high cost of GPU time and specialized supercomputing resources has often kept elite-level drug discovery within the reach of only the largest pharmaceutical conglomerates. By drastically reducing the compute time required to screen billions of compounds, platforms like Ptarmigan-1 make it possible for smaller firms and academic labs to perform massive, "Big Pharma-scale" virtual screens with a fraction of the budget.
As the drug discovery community continues to digest the findings of the Talus Bio preprint, one thing is clear: the era of "structure-only" thinking is ending. The future belongs to models that can navigate the messiness of biological reality—the spaghetti, the disorder, and the motion—rather than those that try to force it into a static, 3D mold. For the millions of patients waiting for therapies for transcription-factor-driven diseases, this shift in computational philosophy may be the most promising development in a decade.
About the Author