NEW YORK, NY & CAMBRIDGE, MA – June 22, 2024 – In a significant leap forward in the fight against one of cancer’s most formidable challenges, researchers at Weill Cornell Medicine and the Massachusetts Institute of Technology (MIT) have uncovered a critical mechanism that appears to dictate whether colorectal cancer (CRC) will spread to the liver. Their collaborative findings point to the loss of GATA6, a vital transcription factor, as a key epigenetic switch that transforms primary tumor cells into highly adaptable, pro-metastatic agents. This discovery, published in the esteemed journal Cell Stem Cell, not only redefines our understanding of metastasis but also paves the way for novel diagnostic and therapeutic strategies to combat this deadliest aspect of colorectal cancer.
Main Facts: A Paradigm Shift in Understanding Metastasis
Colorectal cancer remains a leading cause of cancer-related mortality worldwide, with liver metastasis being the primary driver of patient deaths. For decades, scientists have diligently searched for genetic mutations that trigger this deadly spread, largely with elusive results. This new study introduces a paradigm shift, suggesting that the journey from a localized tumor to a life-threatening metastatic disease may hinge less on DNA sequence alterations and more on profound changes in gene expression orchestrated by epigenetic factors.
The core discovery revolves around GATA6, a transcription factor normally responsible for maintaining the specialized identity and function of intestinal lining cells. The research team found that when GATA6 levels are significantly reduced or lost in colorectal cancer cells, these cells undergo a dramatic transformation. They shed their original identity, entering a more primitive, "fetal-like" and highly adaptable state known as lineage plasticity. This cellular malleability equips them with the extraordinary ability to detach from the primary tumor, survive the perilous journey through the bloodstream, and establish aggressive new tumors in distant organs, most notably the liver.
Crucially, the study demonstrated that this loss of GATA6 is not merely an incidental observation but a causative factor. Its reduction is strongly correlated with poorer patient outcomes and an increased frequency and burden of liver metastases in preclinical models. This groundbreaking insight moves beyond the traditional focus on genetic mutations, highlighting epigenetic reprogramming as a potent force driving cancer dissemination and offering a fresh target for intervention.
Chronology of a Discovery: From Observation to Mechanism
The journey to this pivotal discovery was meticulously structured, beginning with clinical observations and progressing through innovative experimental models to elucidate the underlying molecular mechanisms.
The Metastasis Enigma: A Long-Standing Challenge
For years, the scientific community grappled with the mystery of metastasis. While primary tumor growth is relatively well understood and often treatable, the process by which cancer cells acquire the ability to travel and colonize distant organs has remained a dark horse. Initial hypotheses heavily focused on the accumulation of specific "driver mutations" within the cancer cell’s DNA sequence that would confer metastatic potential. However, despite extensive genomic sequencing efforts, no consistent set of such mutations specifically linked to liver metastasis in CRC had emerged, leaving a critical gap in understanding.
Initial Clues: GATA6 Levels in Metastatic Lesions
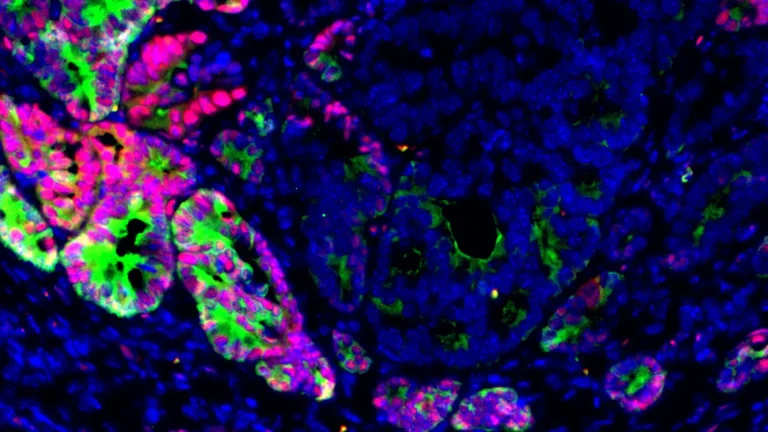
The research began with the observation that GATA6, a transcription factor crucial for intestinal cell identity, exhibited significantly lower levels in liver metastases compared to primary colorectal tumors. This finding was consistent across both mouse models and human patient samples, immediately suggesting a potential link. The correlation between reduced GATA6 expression and poorer patient outcomes further strengthened the hypothesis that GATA6 was not just a bystander but an active participant in the metastatic process.
Developing Novel Models: Capturing Early Metastatic Events
A significant hurdle in metastasis research has been the difficulty in studying the early, transient events that enable cancer cells to become metastatic. As Dr. Norihiro Goto, assistant professor of medicine in the Division of Gastroenterology & Hepatology at Weill Cornell and co-leader of the research, explained, "When researchers analyze patient samples from liver metastases, we fail to capture the important signals occurring in the early stages of the metastatic process." Established metastases represent the end-stage of a complex journey, obscuring the initial transformations.
To overcome this limitation, the team, including Dr. Saori Goto as first author and Dr. Omer H. Yilmaz of MIT as co-leader, developed an ingenious laboratory model using patient-derived organoids. These miniature, three-dimensional clusters of cancer cells, grown in a dish, remarkably recapitulate many biological features of actual tumors. The researchers implanted these organoids into the colons of mice, allowing them to form primary tumors. They then collected cells from the resulting liver metastases and used them to generate new organoids, which were subsequently reimplanted. This iterative process of implantation, metastasis, and organoid derivation allowed the scientists to "evolve" increasingly aggressive, metastatic cancer cells in a controlled environment, providing an unprecedented window into the acquisition of metastatic abilities over time.
Unveiling the Epigenetic Switch: GATA6 and Lineage Plasticity
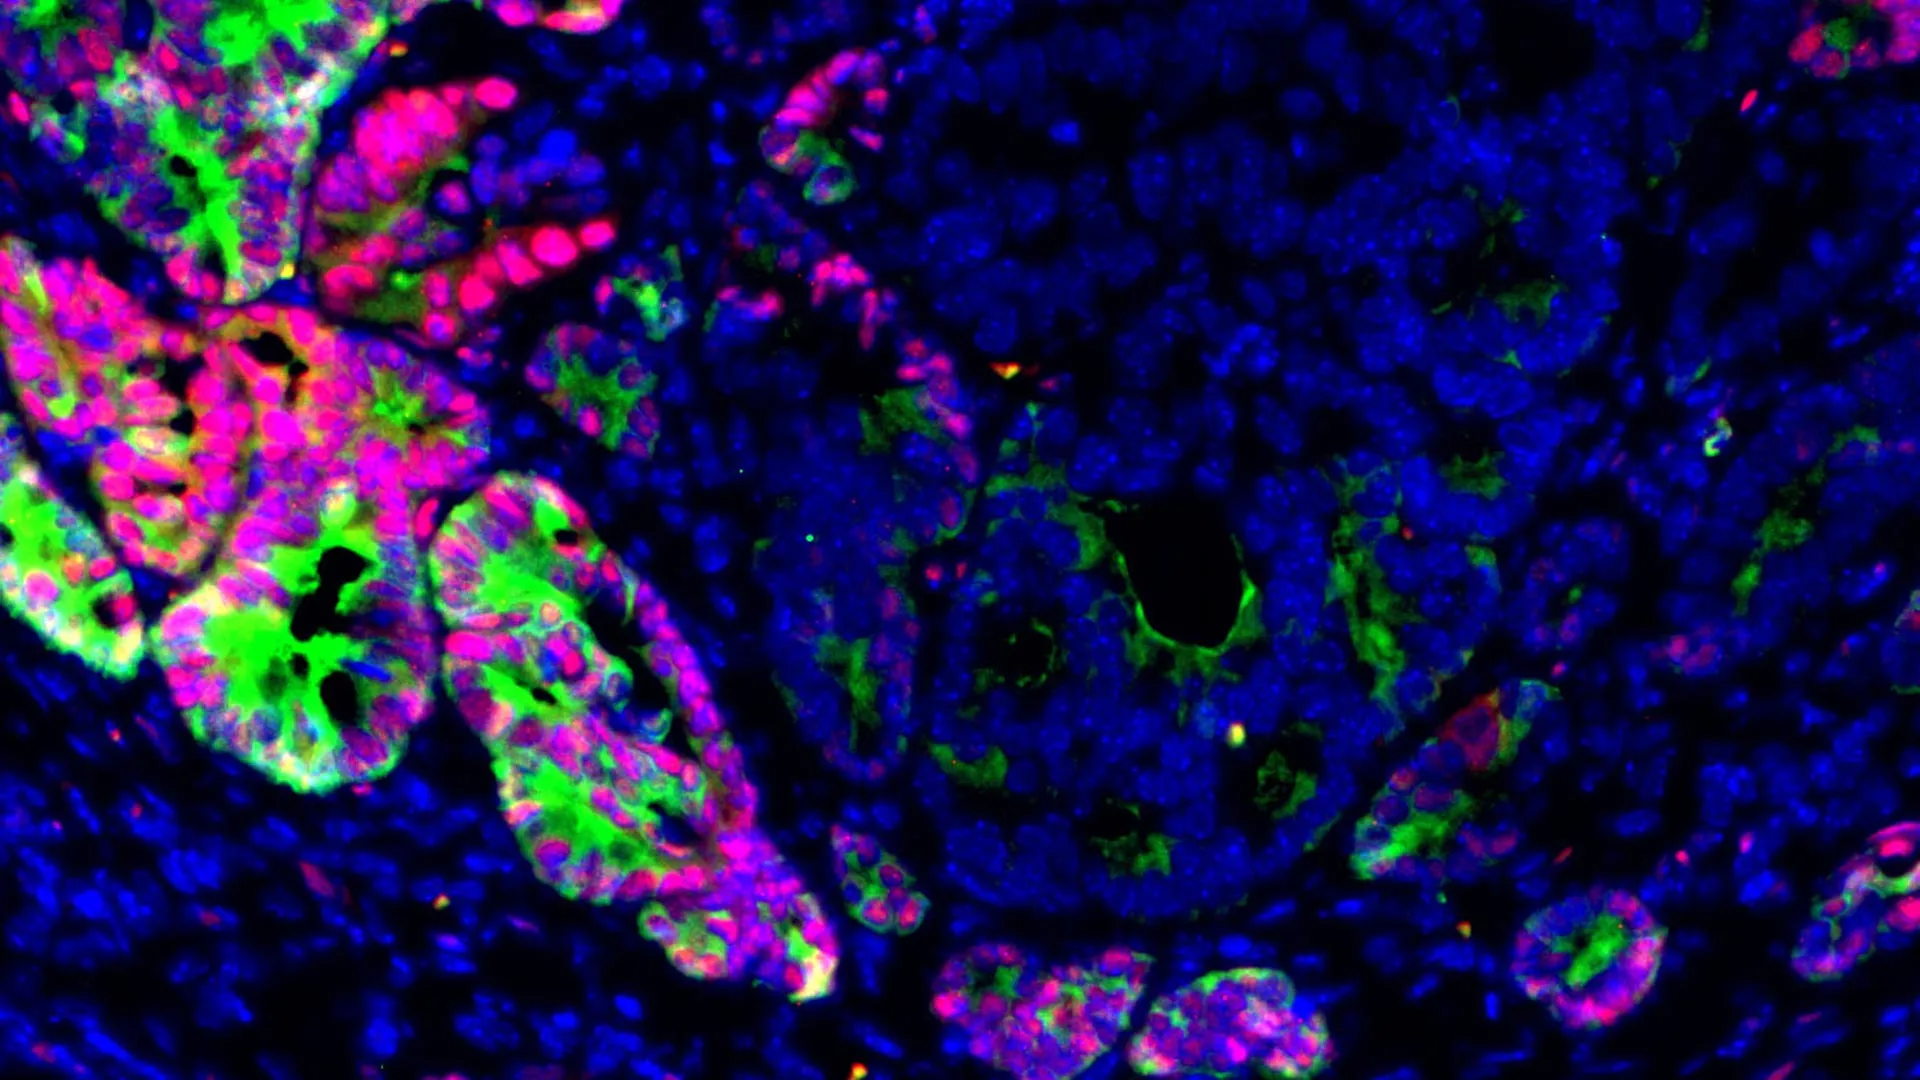
Through these sophisticated organoid models, the researchers meticulously traced the cellular changes associated with increasing metastatic potential. Their experiments unequivocally revealed that the progressive loss of GATA6 was directly responsible for promoting lineage plasticity. This key concept refers to the astonishing ability of cells to alter their identity, function, and behavior. When GATA6 was absent, colorectal cancer cells abandoned their differentiated intestinal cell identity, activating alternative genetic programs. They adopted a flexible, undifferentiated "fetal-like" state – a cellular phenotype normally observed during embryonic development or tissue repair, where cells need to be highly adaptable. This transformation was identified as the critical step that made metastasis possible, allowing these reshaped cells to successfully navigate the hostile environment of the bloodstream and colonize the liver.
Connecting the Dots: LGR5 and Cellular States
Further supporting their findings, the study demonstrated a clear link between GATA6 loss and the expression of LGR5, a well-known marker for intestinal stem cells. Earlier research had suggested that LGR5-negative cells possess a higher capacity to initiate liver metastases. The new study provided the mechanistic explanation: shutting down GATA6 caused a distinct cellular shift from an LGR5-positive to an LGR5-negative state. These LGR5-negative cells, exhibiting fetal-like characteristics, were the ones endowed with enhanced metastatic potential. Conversely, restoring GATA6 activity or activating related pathways effectively reversed this process, significantly reducing the metastatic potential of colorectal cancer cells. This elegant experimental validation solidified GATA6’s role as the central orchestrator of this critical cellular state transition.
Supporting Data: Empirical Evidence and Molecular Insights
The findings are underpinned by a robust body of experimental evidence, integrating molecular biology, pathology, and advanced preclinical models.
Clinical and Preclinical Correlation:
- Human Patient Data: Analysis of human liver metastases from CRC patients consistently showed significantly lower levels of GATA6 compared to primary colorectal tumors. This observation was crucial, as it linked the laboratory findings directly to the clinical reality of the disease.
- Patient Outcomes: Reduced GATA6 expression in patient tumors was directly associated with poorer prognosis and shorter survival times, underscoring its clinical relevance as a prognostic indicator.
- Mouse Models: In genetically engineered mouse models of colorectal cancer, the deliberate genetic deletion of GATA6 dramatically increased both the frequency and burden of liver metastases. Importantly, this deletion had "little effect on primary tumor growth," as noted by Dr. Norihiro Goto. This specific impact on metastasis, rather than primary tumor bulk, highlights that GATA6 loss drives a distinct metastatic program independent of initial tumor proliferation.
Organoid Model Validation:
- Iterative Selection: The innovative organoid model, involving serial transplantation and re-derivation, proved instrumental. Each successive generation of organoids derived from liver metastases exhibited progressively lower GATA6 levels and a greater capacity for metastasis, directly demonstrating the selective pressure for GATA6 loss in metastatic progression.
- Molecular Characterization: Gene expression profiling of GATA6-deficient organoids and metastatic cells revealed activation of alternative genetic programs characteristic of a more primitive, undifferentiated state. These programs included genes associated with epithelial-mesenchymal transition (EMT) – a process known to confer migratory and invasive properties – and stemness, both hallmarks of enhanced metastatic potential.
Mechanistic Elucidation of Lineage Plasticity:
- LGR5 Status Shift: Detailed cellular analysis confirmed that GATA6 loss directly induced a phenotypic switch from LGR5-positive to LGR5-negative cells. This transition was accompanied by a re-expression of fetal-like genes and a heightened ability to form metastatic colonies in secondary sites.
- Restoration Experiments: Crucially, the researchers demonstrated causality by reintroducing GATA6 into GATA6-deficient metastatic cells. This restoration led to a partial re-differentiation of the cells, a reduction in their plastic state, and a significant decrease in their ability to metastasize in experimental models. This "rescue" experiment provided strong evidence that GATA6 loss is not merely correlative but functionally critical for promoting metastasis.
- Epigenetic Signatures: While the study primarily focused on the functional outcome of GATA6 loss, the researchers also delved into the underlying epigenetic changes. They observed alterations in chromatin accessibility and histone modifications associated with the activation of alternative gene regulatory networks when GATA6 was absent, confirming the epigenetic nature of the observed cellular reprogramming.
These comprehensive data collectively paint a clear picture: GATA6 acts as a crucial epigenetic gatekeeper, and its loss unlocks a potent metastatic program driven by cellular plasticity, rather than solely by new genetic mutations.
Official Responses: Expert Insights and Collaborative Vision
The researchers involved in this landmark study articulated the significance of their findings and the collaborative spirit that drove the discovery.
Dr. Norihiro Goto, assistant professor of medicine in the Division of Gastroenterology & Hepatology at Weill Cornell Medicine and co-leader of the research, emphasized the core finding: "We discovered that GATA6 loss acts as a critical switch that can change cancer cells in the primary tumor from non-metastatic to pro-metastatic." His statement underscores the transformative power of this epigenetic event, shifting the cellular identity and functional capabilities of the cancer cells. He further highlighted the broader implications for cancer research, noting, "Our findings suggest that epigenetic changes may be more important for promoting liver metastasis." This statement directly challenges the long-held mutation-centric view of metastasis, advocating for a more nuanced understanding of cellular reprogramming.
Dr. Goto also reflected on the methodological innovation that enabled this discovery. "When researchers analyze patient samples from liver metastases, we fail to capture the important signals occurring in the early stages of the metastatic process," he explained, justifying the team’s development of sophisticated organoid models. These models allowed them to observe the dynamic acquisition of metastatic traits, a crucial step that traditional biopsy analysis often misses. He also added, "When we genetically delete GATA6, the frequency and burden of liver metastases in mouse models significantly increase, while having little effect on primary tumor growth," providing clear evidence of GATA6’s specific role in metastasis rather than primary tumor expansion. As a member of the Jill Roberts Institute for Research in Inflammatory Bowel Disease and Sandra and Edward Meyer Cancer Center, both at Weill Cornell, Dr. Goto’s insights bridge fundamental research with clinical relevance.
Dr. Saori Goto, an instructor in medicine at Weill Cornell and first author of the study, played a crucial role in meticulously executing the experimental work that led to these insights. Her efforts in characterizing the epigenetic landscape and cellular transformations driven by GATA6 loss were instrumental in building the robust evidence base for the paper.
Dr. Omer H. Yilmaz, associate professor of biology at the Massachusetts Institute of Technology, who also co-led the work, contributed his expertise in stem cell biology and organoid technology, which was critical for developing the innovative preclinical models used in the study. His involvement underscores the interdisciplinary nature of modern biomedical research, bringing together diverse scientific strengths to tackle complex biological problems.
Collectively, the researchers’ statements convey a clear message: this study represents a significant departure from previous approaches to understanding metastasis, offering a fresh perspective rooted in epigenetic regulation and cellular plasticity. Their collaborative vision extends beyond discovery, aiming to translate these insights into tangible benefits for patients. Dr. Norihiro Goto concluded, "In addition to treating primary tumors, we need to find strategies to target the mechanism of liver metastasis. Our study is a step toward developing therapies that block the spread of cancer at the earliest stages." This forward-looking perspective encapsulates the ultimate goal: to move from understanding to intervention, ultimately saving lives.
Implications: Biomarkers, Therapies, and Future Directions
The implications of this groundbreaking research are far-reaching, promising to reshape how colorectal cancer is diagnosed, monitored, and treated, particularly concerning its metastatic spread.
1. Potential Biomarker for Metastatic Risk:
One of the most immediate and impactful implications is the potential for GATA6 to serve as a novel biomarker. Currently, predicting which primary colorectal tumors will metastasize to the liver remains challenging. The study suggests that tumors exhibiting low levels of GATA6 expression may harbor a greater proportion of cells capable of undergoing the metastasis-promoting cellular state transition.
- Patient Stratification: This could enable clinicians to identify patients at higher risk of developing liver metastases much earlier in their disease course. Such identification would allow for more personalized and aggressive treatment strategies, including closer monitoring, adjuvant therapies, or even prophylactic interventions before overt metastasis occurs.
- Prognostic Value: GATA6 levels could become a standard part of diagnostic pathology, providing crucial prognostic information that guides treatment decisions and patient counseling.
- Liquid Biopsies: Future research might explore GATA6 expression or related epigenetic signatures in circulating tumor DNA (ctDNA) or circulating tumor cells (CTCs) as a non-invasive tool for real-time monitoring of metastatic potential.
2. Novel Therapeutic Strategies Targeting Cellular Identity:
The findings open an entirely new therapeutic avenue: maintaining cellular identity and preventing cancer cells from entering highly flexible, pro-metastatic states. Instead of solely focusing on killing rapidly dividing cells (the basis of traditional chemotherapy), future therapies could aim to:
- Restore GATA6 Activity: Developing small molecules or gene therapies that can restore GATA6 expression or activate its downstream pathways in cancer cells could suppress their lineage plasticity and reduce metastatic potential.
- Inhibit Plasticity Pathways: Identifying and targeting the specific alternative genetic programs activated when GATA6 is lost could prevent cells from adopting the fetal-like metastatic state. This might involve inhibiting transcription factors or signaling pathways that become hyperactive in GATA6-deficient cells.
- "Identity Lock-In" Therapies: The concept of "identity lock-in" involves pharmacologically reinforcing the differentiated state of cancer cells, making them less capable of adaptation and spread. This approach could render metastatic cells less aggressive and more susceptible to existing treatments.
- Challenges: Dr. Norihiro Goto rightly cautioned that researchers will need to find ways to target these processes without interfering with normal tissue repair, which also relies on similar biological programs of cellular plasticity. Therapeutic specificity will be paramount to avoid unwanted side effects.
3. Rethinking Primary Tumor Growth vs. Metastasis:
The observation that GATA6 loss significantly impacts metastasis without substantially affecting primary tumor growth represents a crucial conceptual shift. It suggests that the mechanisms driving primary tumor proliferation are distinct from those enabling metastatic dissemination. This calls for a dual therapeutic approach: one targeting the primary tumor and another specifically designed to block or reverse the metastatic cascade.
4. Future Research Directions:
The study lays a robust foundation for extensive future research:
- Identifying Vulnerabilities: A key next step will be to identify unique vulnerabilities specific to GATA6-deficient, plastic cancer cells. These vulnerabilities could represent novel therapeutic targets that are less likely to affect normal, differentiated cells. This might involve exploring metabolic dependencies, signaling pathway alterations, or immunogenic changes in these transformed cells.
- Tumor Microenvironment Influence: The team plans to investigate how the tumor microenvironment, including immune cells, fibroblasts, and liver-specific signals, influences these cellular transitions in preclinical models. The liver microenvironment plays a critical role in supporting metastatic growth, and understanding its interaction with plastic cancer cells could reveal additional therapeutic targets.
- Combination Therapies: Exploring combination therapies that target GATA6-driven plasticity alongside conventional treatments (e.g., chemotherapy, immunotherapy) could lead to synergistic effects, improving patient outcomes.
- Translational Studies: Moving from preclinical models to human clinical trials will be essential to validate GATA6 as a biomarker and test the efficacy and safety of plasticity-modulating therapies.
In summary, this research not only provides a powerful new understanding of colorectal cancer metastasis but also offers tangible pathways for developing innovative strategies to prevent its deadliest manifestation. By shifting focus to epigenetic regulation and cellular identity, the scientific community gains a potent new weapon in the ongoing battle against advanced cancer. The ultimate goal, as articulated by Dr. Norihiro Goto, is to "block the spread of cancer at the earliest stages," transforming the prognosis for countless patients worldwide.
This research was supported in part by the Astellas Foundation; Research Abroad from Japan Society for the Promotion of Science; the National Institutes of Health (grants R00AG076987, 01CA254314,5U01CA25055, R01CA258523, R01CA25723, R01DK133919, R01DK140310, R01CA299955, and 3OT2CA297570); Pew-Stewart Trust; AFAR and Glenn Foundation for Medical Research Breakthroughs in Gerontology; Kenneth Rainin Foundation; Crohn’s & Colitis Foundation and Mark Foundation for Cancer Research.
About the Author