The pharmaceutical landscape underwent a seismic shift in May 2026 when the U.S. Food and Drug Administration (FDA) granted approval to vepdegestrant (Veppanu®). As the first-ever Proteolysis Targeting Chimera (PROTAC) to reach the commercial market, vepdegestrant represents more than just a new therapeutic for ER-positive, HER2-negative, ESR1-mutated breast cancer; it serves as a historic proof-of-concept for the entire modality of targeted protein degradation (TPD).
For years, TPD was relegated to the realm of high-potential, high-risk academic inquiry. Today, it stands as a clinically validated approach. However, the path from bench to bedside remains fraught with technical hurdles, the most significant of which is the transition from parenteral administration to the patient-preferred oral route. As drug developers look to replicate the success of vepdegestrant, they must grapple with the fundamental "oral exposure problem"—a multifaceted challenge that requires a departure from traditional medicinal chemistry rules.
The Mechanism: Redefining "Druggability"
At the heart of the PROTAC revolution is a departure from the "occupancy-driven" paradigm that has defined small-molecule pharmacology for decades. Traditional inhibitors must bind to a protein’s active site with high affinity and maintain that occupancy to achieve a therapeutic effect. PROTACs, conversely, operate on an "event-driven" catalytic mechanism.
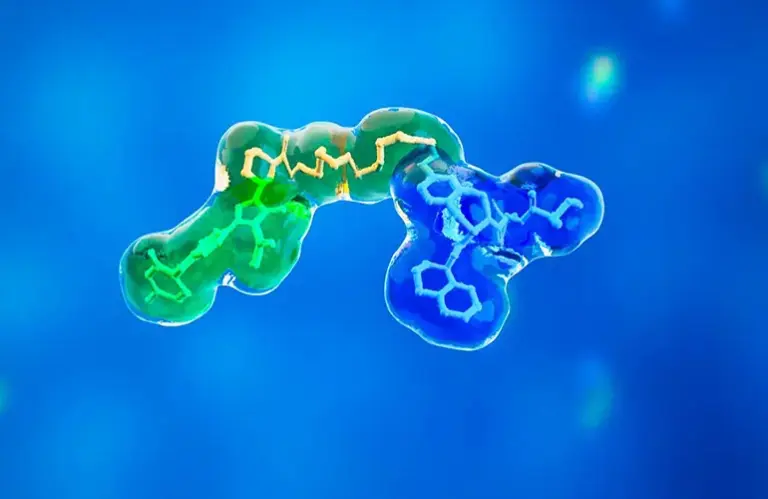
A typical PROTAC molecule is a heterobifunctional construct comprising three key elements: a warhead that binds the target protein, a ligand that recruits an E3 ubiquitin ligase, and a linker that tethers the two. By bringing the target protein into proximity with an E3 ligase, the PROTAC induces polyubiquitination, marking the target for destruction by the cell’s internal waste-disposal machinery—the proteasome.
This mechanism offers profound clinical advantages. Because the PROTAC acts as a catalyst rather than a stoichiometric blocker, it does not need to saturate the target protein; a single molecule can facilitate the degradation of multiple copies of the target. This enables efficacy at lower concentrations and provides a pathway to target "undruggable" proteins that lack clear binding pockets for traditional inhibitors. Yet, these very biological benefits—molecular size, complexity, and specialized chemical architecture—create substantial barriers to oral absorption.
Chronology of a Modality: From Concept to Clinical Reality
The evolution of PROTACs has been a study in iterative engineering. While the fundamental concept was proposed in the early 2000s, the field spent the subsequent two decades refining the structural components of these degraders.
- 2001–2010: The Conceptual Foundation. Initial studies proved the feasibility of using chimeric molecules to bridge target proteins and E3 ligases. However, these early iterations were largely cellular probes with poor pharmacokinetic profiles.
- 2011–2018: Optimization and Expansion. The identification of potent ligands for E3 ligases like Cereblon (CRBN) and Von Hippel-Lindau (VHL) catalyzed the field. Developers began moving beyond simple proof-of-concept studies toward optimized, drug-like candidates.
- 2019–2025: The Preclinical Surge. As the industry raced to solve the "beyond-Rule-of-Five" (bRo5) problem, massive investment poured into DMPK (Drug Metabolism and Pharmacokinetics) and translational science to predict human oral exposure from animal models.
- May 2026: Regulatory Validation. The FDA approval of vepdegestrant marked the official "coming of age" for PROTACs. The approval demonstrated that, with the right optimization strategies, these complex molecules could indeed be formulated as successful oral medicines.
Supporting Data: The "Beyond-Rule-of-Five" Hurdle
The primary obstacle in PROTAC development is that they frequently violate Lipinski’s Rule of Five (Ro5). Traditional small molecules are typically below 500 Da with moderate polarity. PROTACs, by necessity of their bifunctional design, often exceed 700 Da, feature high polar surface areas (>150 Ų), and contain more than 15 rotatable bonds.
According to data analyzed in the field, these characteristics lead to poor membrane permeability and suboptimal aqueous solubility. Traditional screening methods that rely on Ro5 parameters often flag these candidates as "non-oral." However, recent research suggests that Ro5 is a poor predictor for PROTACs.
A critical 2024 study published in the Journal of Medicinal Chemistry highlighted that the number of solvent-exposed hydrogen-bond donors (eHBD) is a far more reliable discriminator for oral bioavailability. The study suggests an upper limit of approximately eHBD ≤ 2 for oral candidates. This finding has reframed how lead optimization is conducted, shifting the focus from simple weight reduction to the strategic management of molecular polarity and conformational folding.
Official Industry Perspectives: The Strategic Blueprint
Industry leaders, including Tao Xiong of WuXi AppTec, emphasize that solving the oral exposure challenge requires a cross-functional approach that begins at the earliest stages of discovery.
1. The Role of the Food Effect
One of the most practical, yet often overlooked, factors in PROTAC oral bioavailability is the "food effect." Because many PROTACs have solubility-limited absorption, their performance in the gastrointestinal tract can vary drastically between fasted and fed states. Vepdegestrant’s label, which recommends administration with food, is a prime example of this strategy in action. Developers are now utilizing biorelevant intestinal media (e.g., FaSSIF and FeSSIF) to mimic human conditions, ensuring that candidate selection is based on data that reflects real-world clinical performance.
2. Linker Engineering and Permeability
The linker is not merely a spacer; it is a fundamental determinant of the molecule’s ability to cross biological membranes. Replacing flexible PEG chains with rigid structures, such as 1,4-disubstituted phenyl rings, has been shown to reduce molecular flexibility and improve permeability. Furthermore, reducing amide motifs and managing the nitrogen content within the linker can help the molecule adopt "folded" conformations that shield polar groups, effectively "hiding" the molecule from the aqueous environment while it traverses lipid bilayers.
3. E3 Ligase Selection as a Developability Strategy
The choice of E3 ligase is as much a pharmacokinetic decision as a biological one. While VHL and CRBN are the industry standards, CRBN-based degraders often allow for more compact molecular architectures. This compactness is a significant advantage in the quest for oral bioavailability. When selecting a ligase, teams must weigh the protein-degradation efficiency against the structural "bulk" the ligase ligand adds to the overall construct.
4. The Prodrug Paradigm
When structural optimization hits a ceiling, prodrug strategies offer a potential bypass. By modifying an active degrader to be more lipophilic or less polar, developers can enhance absorption, relying on in vivo enzymatic conversion to release the active drug. While this adds complexity to the regulatory and manufacturing process, the success of lipid-based or masked-ligand prodrugs in preclinical trials suggests this will be a cornerstone of future TPD development.
5. Embracing Alternatives: Molecular Glues
It is important to recognize that PROTACs are not a panacea. When a target is particularly stubborn or the PROTAC structure becomes prohibitively large, "molecular glues"—smaller molecules that stabilize protein-protein interactions without a linker—may offer a more viable path to an oral drug. Molecular glues provide a simpler starting point for permeability and systemic exposure, serving as a critical alternative tool in the pharmaceutical arsenal.
Implications for the Future of Drug Discovery
The successful approval of vepdegestrant has effectively "opened the floodgates" for TPD therapeutics. However, the industry is now entering a phase of "sophisticated development." The era of simply creating a potent degrader in a lab is over; the current era demands the creation of a drug.
The implications for developers are clear:
- Integrated DMPK: ADME (Absorption, Distribution, Metabolism, and Excretion) studies cannot be an afterthought. They must be integrated into the medicinal chemistry design cycle.
- Early Bioanalysis: The use of ultrasensitive bioanalytical platforms is now essential to detect and quantify these complex molecules in systemic circulation at very low doses.
- Cross-Functional Collaboration: The silos between structural biologists, chemists, and pharmacologists must be dismantled. Oral PROTAC optimization is a team sport where structural biology informs conformational design, which in turn is validated by fit-for-purpose cell-based assays.
As we look toward 2030, the ability to turn PROTACs into oral medicines will define the next generation of oncology and beyond. By treating oral bioavailability as a multifaceted challenge—encompassing structural biology, biorelevant testing, and deliberate molecular engineering—the biopharmaceutical industry is poised to move beyond the "undruggable" barriers of the past and into a new age of precision, orally administered protein degradation.
The success of the first generation of PROTACs is not the end of the journey; it is the calibration point for a new standard in drug development. For those capable of navigating the complex nexus of chemistry, biology, and pharmacokinetics, the rewards in patient outcomes and therapeutic impact will be substantial.
About the Author