In a significant stride toward transforming the landscape of vascular medicine, Teleflex Incorporated has announced that its Freesolve Resorbable Magnesium Scaffold (RMS) has been granted Breakthrough Device Designation by the U.S. Food and Drug Administration (FDA). This regulatory milestone, specifically targeting the device’s application in percutaneous coronary intervention (PCI), highlights the growing momentum behind "leave-nothing-behind" technologies. By replacing traditional permanent metallic stents with a temporary, bioresorbable magnesium-based structure, Teleflex aims to address the limitations of conventional coronary implants and offer a more physiological approach to vessel healing.
The Core Innovation: Engineering a Temporary Scaffold
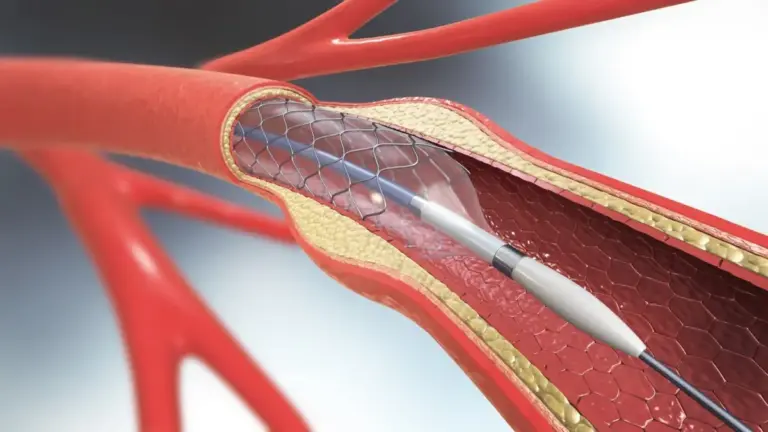
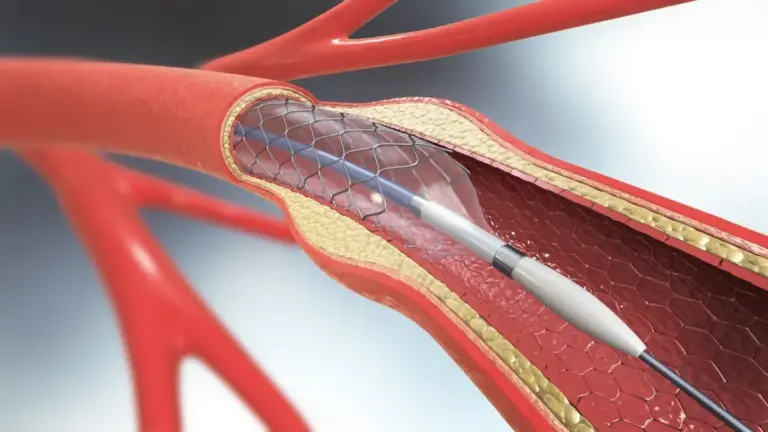
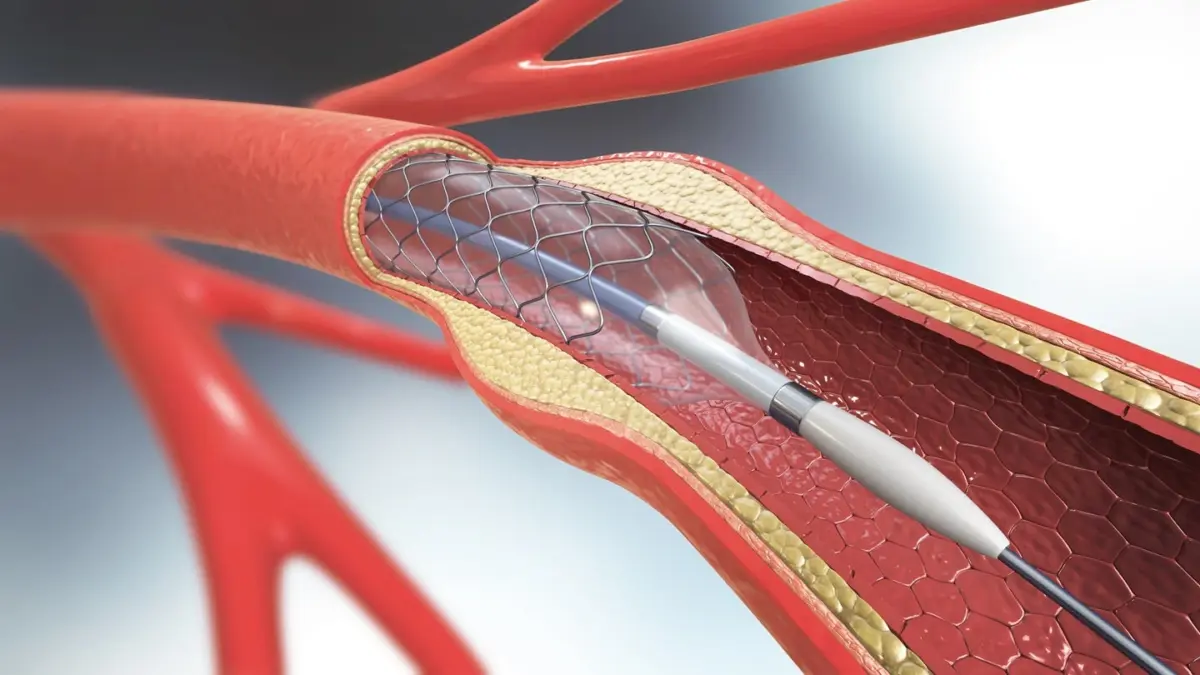
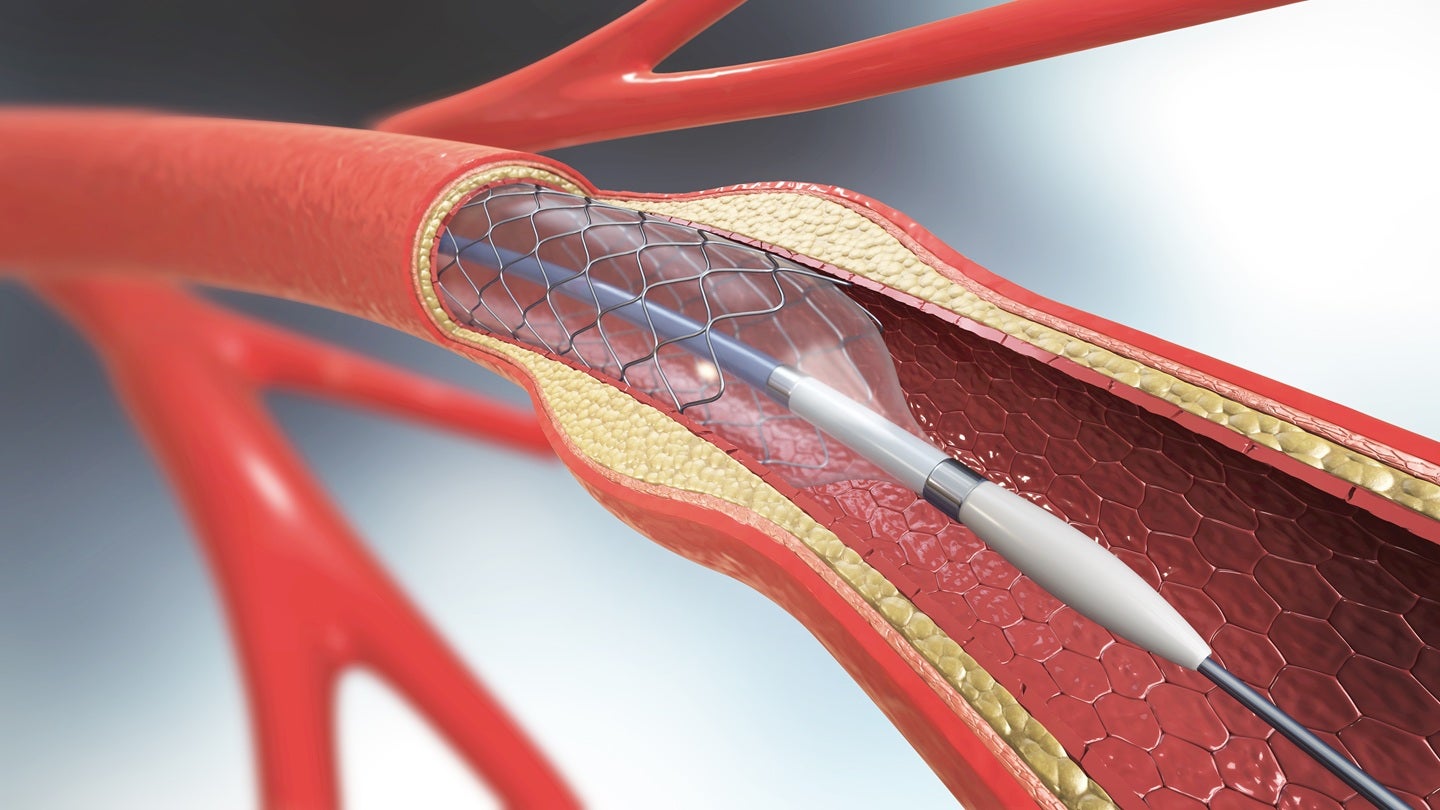
At the heart of the Freesolve RMS lies a sophisticated marriage of material science and clinical engineering. The device is constructed from a proprietary BIOmag magnesium alloy, which provides the necessary radial strength to support the coronary artery during the critical period following an angioplasty. Unlike permanent stents—which remain in the body for the patient’s lifetime and can pose long-term risks such as late-stent thrombosis, neoatherosclerosis, or the limitation of future surgical bypass options—the Freesolve scaffold is engineered to be temporary.
The scaffold is enhanced with BIOlute Bioabsorbable Coating Technology, which ensures the controlled delivery of therapeutic agents to prevent restenosis while the vessel undergoes the remodeling process. According to the company, the body resorbs approximately 99% of the magnesium scaffold within a 12-month window. Once the scaffold is absorbed, the vessel is left free of foreign materials, potentially restoring natural vasomotion and allowing the artery to return to a more natural, healthy state.
Chronology: From Concept to Breakthrough
The journey of the Freesolve RMS is a testament to the rigorous, multi-year validation process required to bring disruptive medical technology to the U.S. market.
- Early Clinical Validation: The journey began with the BIOMAG-I First-In-Human (FIH) study. This trial provided the initial evidence that magnesium-based scaffolding could provide stable support without the long-term inflammatory risks associated with permanent steel or cobalt-chromium cages.
- International Expansion: Before seeking U.S. approval, Teleflex pursued and secured the CE Mark, allowing for commercialization in the Asia-Pacific region, Europe, the Middle East, and South America. This global presence provided real-world data that served as a foundation for subsequent regulatory filings.
- The Below-the-Knee Milestone: In 2024, the FDA granted Breakthrough Device Designation to the Freesolve scaffold for below-the-knee (BTK) applications, specifically for patients suffering from chronic limb-threatening ischemia (CLTI). This was a critical precursor, signaling the FDA’s willingness to embrace the magnesium alloy technology.
- The Current Phase: Today, Teleflex is actively conducting the BIOMAG-III Pivotal Trial. This large-scale study is designed to generate the robust clinical data necessary to support a Premarket Approval (PMA) application, which will be the final hurdle for full FDA clearance in the coronary space.
Supporting Data: Evidence of Long-Term Efficacy
The clinical enthusiasm for Freesolve is rooted in the long-term follow-up data from the BIOMAG-I study. In the world of interventional cardiology, the "four-year mark" is a significant threshold for assessing the durability of a vascular intervention.
The BIOMAG-I trial revealed a 3.5% rate of target lesion failure (TLF) at the four-year follow-up interval. Perhaps more importantly, the study reported no new clinical events between the second and fourth years post-implantation. This absence of late-term clinical "rebound" is a major selling point for the technology, as it suggests that the transition from a scaffolded vessel to an unencumbered native vessel occurs smoothly and without late-stage complications.
The ongoing BIOMAG-III Pivotal Trial is currently expanding upon this, utilizing a broader patient population to ensure that the findings of the initial FIH studies hold true across the diverse patient profiles seen in modern catheterization labs.
Official Perspectives: A Strategic Commitment
The leadership at Teleflex views the Breakthrough Device Designation as more than just a regulatory win; they see it as validation of their long-term strategic pivot toward advanced vascular solutions.
Jason Fontana, President and Interventional General Manager at Teleflex, emphasized the company’s vision in a recent statement: "At Teleflex, we are advancing vascular intervention through strategic investment and continuous innovation. This designation marks an important milestone in our efforts to bring Freesolve RMS to patients in the U.S."
Fontana further underscored the synergy between the current coronary application and the previous BTK designation, noting, "Building on the FDA’s previous breakthrough device designation for Freesolve RMS in below-the-knee applications, it further underscores the potential of bioresorbable magnesium scaffold technology and our commitment to a leave-nothing-behind approach. Freesolve RMS has the potential to redefine clinical practice in today’s cath labs."
The FDA’s Breakthrough Device Program is designed to expedite the regulatory review of technologies that offer significant advantages over existing approved alternatives for life-threatening or irreversibly debilitating conditions. By granting this status, the FDA has acknowledged that Freesolve could provide more effective treatment for patients who might otherwise require lifelong medication or permanent implants.
Implications for Clinical Practice and Patient Care
The potential for Freesolve to "redefine clinical practice" carries profound implications for both the interventionalist and the patient.
For the Interventionalist
The current standard of care—permanent drug-eluting stents (DES)—has been remarkably successful, yet it is not without drawbacks. Physicians are often tasked with managing the long-term implications of permanent metal cages, such as the potential for late-stage thrombosis or the inability to re-access a vessel for future bypass surgery. Freesolve offers a "reset" button. If the scaffold is absorbed, the coronary artery is theoretically preserved for future interventions, a benefit that cannot be overstated in a patient population that is living longer and potentially requires multiple procedures over a lifetime.
For the Patient
Patients are increasingly seeking treatments that prioritize long-term quality of life over the mere immediate mechanical opening of a vessel. The idea of an "implant-free" vessel following a one-year healing window is highly appealing. By minimizing the amount of foreign material left in the body, the Freesolve technology aims to reduce the patient’s long-term dependence on dual-antiplatelet therapy (DAPT), potentially improving long-term health outcomes and reducing the risk of bleeding complications associated with extended blood-thinning regimens.
Market Dynamics and Future Outlook
The move by Teleflex places it in a competitive position within the high-stakes global market for coronary interventions. While the "bioresorbable" category has faced historical challenges—most notably the market withdrawal of previous-generation scaffolds from other manufacturers—the industry has learned valuable lessons regarding strut thickness, material degradation rates, and patient selection.
Teleflex’s use of magnesium, as opposed to the polymers used in earlier iterations of bioresorbable scaffolds, is seen by many in the industry as a technological evolution. Magnesium possesses physical properties that allow for thinner struts—which improves deliverability and reduces vessel trauma—while maintaining the structural integrity required to prevent vessel recoil.
As the BIOMAG-III trial progresses, the industry will be watching closely to see if the success seen in the BIOMAG-I data can be replicated at scale. Should the trial meet its endpoints, the Freesolve RMS could very well become a centerpiece of the modern cath lab, signaling a definitive shift toward a future where "permanent" is no longer the default for coronary intervention.
In conclusion, the FDA’s Breakthrough Device Designation for the Freesolve RMS represents a critical juncture for Teleflex. It is a validation of the technical merits of their magnesium-alloy platform and a clear signal that the regulatory environment is increasingly supportive of technologies that challenge the status quo. As Teleflex continues to navigate the path toward full PMA approval, the medical community waits in anticipation for a tool that promises not just to treat the disease, but to eventually fade away, leaving the patient’s vessel as close to its natural state as possible.
About the Author