In a landmark decision for genomic medicine, the U.S. Food and Drug Administration (FDA) has granted accelerated approval to Inovio Pharmaceuticals for Voceryq (doruxapapogene ralaplasmid and lalikinogene sifuplasmid-gxfd). As the first plasmid DNA-based immunotherapy approved for human use, Voceryq represents a significant technological leap in treating recurrent respiratory papillomatosis (RRP). However, the regulatory victory comes with strings attached: the FDA has mandated that Inovio conduct rigorous postmarketing reevaluations of the Cellectra 5PSP—the electroporation delivery device essential to the drug’s administration.
While Inovio prepares for a commercial rollout, the agency’s requirement for further human factors testing and sharps injury prevention verification highlights the ongoing tension between pioneering complex delivery hardware and ensuring absolute clinical safety in real-world settings.
The Regulatory Landscape: Approval with Conditions
On October 30, 2026, the FDA provided a conditional green light for Voceryq, a therapeutic designed to address the debilitating symptoms of RRP—a rare condition where human papillomavirus (HPV) triggers chronic, recurring growths in the airway. Because standard treatment typically involves repetitive, invasive surgeries, Voceryq’s promise to reduce the surgical burden is viewed as a major clinical advancement.
Despite the approval, the FDA’s letter to Inovio underscores a cautious approach toward the Cellectra 5PSP device. The agency has issued two primary postmarketing requirements (PMRs) that necessitate immediate attention:
- Human Factors Reanalysis: Inovio must re-examine its human factors validation study. This is not a request for a simple review, but a demand to granularly separate "close calls" from "use difficulties." The company is tasked with performing a comprehensive root-cause and clinical risk analysis to ensure that healthcare providers—who must be specially trained to operate the device—can do so without error.
- Sharps Injury Prevention: The FDA has mandated formal verification and validation testing of the device’s sharps injury prevention features. Given that the device involves an injection sequence followed by electrical pulses, the potential for accidental needle sticks is a critical safety parameter that the regulator wants further documented.
Inovio has until June 30, 2027, to submit final reports. Should the analysis suggest that the device’s current design is insufficient, the company will be required to submit proposed modifications and supplemental validation protocols by that date, followed by a formal prior-approval supplement by the end of 2027.
Understanding the Technology: DNA and Electroporation
Voceryq is not a traditional vaccine or pharmaceutical; it is a plasmid DNA-based immunotherapy. Plasmid DNA consists of small, circular genetic loops that carry instructions to produce specific proteins. When injected into muscle tissue, these instructions teach the patient’s T cells to recognize and eradicate HPV-infected papilloma cells.
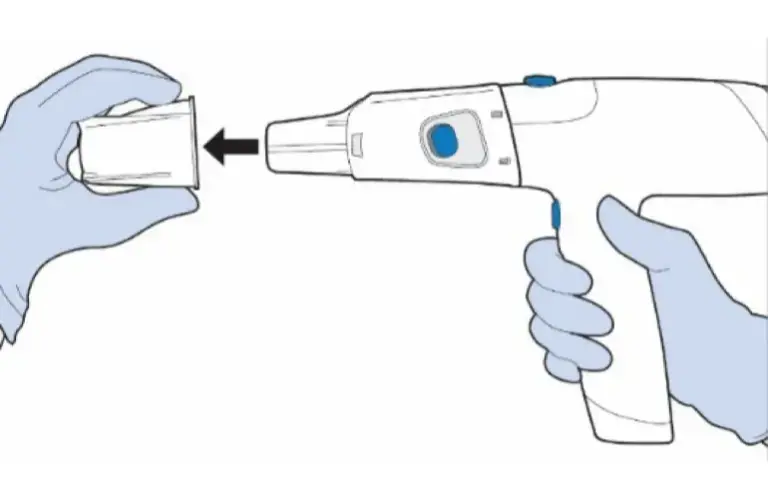
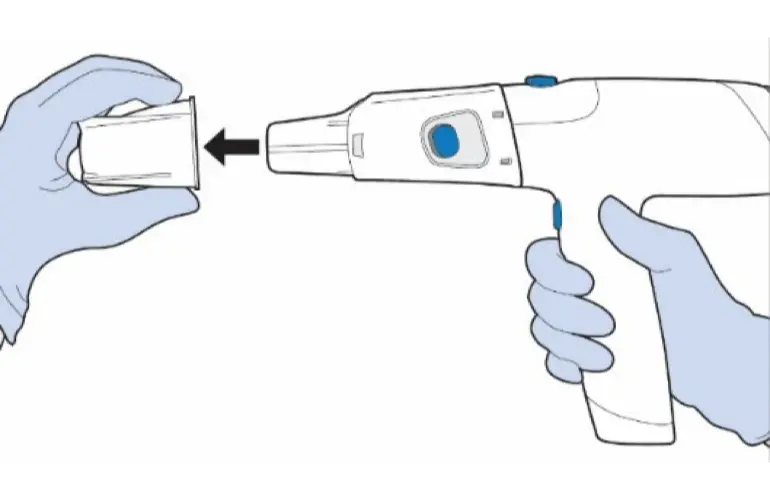
The efficiency of this process hinges entirely on the Cellectra 5PSP device. DNA molecules are large and struggle to cross the protective membranes of human cells on their own. The Cellectra 5PSP uses a process called electroporation, which delivers precise, brief electrical pulses to the tissue. These pulses create temporary pores in the cell membranes, allowing the DNA plasmids to enter and begin their work.
The administration protocol is exacting. Once triggered, the delivery sequence takes approximately 10 seconds. Following the surgical removal of visible growths, patients are prescribed a 1 mL dose administered every three weeks for a four-dose regimen. Furthermore, the FDA label requires at least 30 minutes of post-treatment observation to monitor for local site reactions, reflecting the potent nature of the electroporation-facilitated delivery.
Chronology of Inovio’s Regulatory Journey
Inovio’s history with the FDA has been marked by periodic friction regarding its delivery hardware, suggesting that the current requirements for Voceryq are part of a long-standing regulatory dialogue.
- October 2016: The FDA placed a clinical hold on the Phase 3 program for VGX-3100, another of Inovio’s DNA immunotherapies. The hold was prompted by concerns regarding the shelf-life data of the disposable components of the Cellectra device. At the time, the trial was in its infancy, and the hold effectively paused enrollment.
- June 2017: After providing the necessary documentation and data to satisfy the agency’s concerns regarding device reliability, Inovio successfully had the hold lifted, allowing the VGX-3100 program to resume.
- August 2026: The FDA updated its formal guidance on "Applying Human Factors and Usability Engineering to Medical Devices." This updated framework set the bar higher for how manufacturers document "close calls"—instances where a user makes an error but recovers before harm occurs—and "difficulties," such as confusion during a procedure.
- October 30, 2026: The FDA grants accelerated approval for Voceryq, explicitly linking the drug’s availability to the Cellectra 5PSP and mandating the aforementioned postmarketing studies.
Supporting Clinical Evidence
The approval of Voceryq is predicated on data from an open-label, single-arm clinical trial involving 32 adult patients. The primary efficacy endpoint was a reduction in the frequency of surgeries, which the FDA accepted as a surrogate endpoint "reasonably likely to predict clinical benefit."
In the FDA’s analysis of 26 patients, 27% (seven individuals) remained surgery-free for a full year following the treatment. These patients were specifically selected for their high disease burden, having undergone at least three surgeries in the 12 months preceding their participation in the trial. A secondary, retrospective follow-up study conducted by the investigators found that 14 of 28 participants required no surgery in the second year, suggesting a potential for durable therapeutic effect.
However, the "accelerated" nature of this approval means the burden of proof remains high. Inovio is currently obligated to conduct a confirmatory study to verify these long-term benefits. The final report for this confirmatory trial is due by August 30, 2032. Failure to provide sufficient evidence of efficacy or a failure to pursue this research with "due diligence" could result in the FDA withdrawing approval for the drug entirely.
Official Responses and Strategic Implications
Inovio has publicly downplayed the impact of the FDA’s postmarketing requirements on its commercial timeline. In a statement provided to Drug Discovery and Development, an Inovio spokesperson noted that postmarketing commitments are a standard component of the FDA approval process for innovative technologies.
"We look forward to working with the agency to complete [the requirements]," the spokesperson stated. "We do not expect them to affect our plans to make Voceryq available to order and ship in approximately 60 days, with full field deployment in the first quarter of 2027."
Implications for the Rare Disease Market
The approval of Voceryq marks a milestone for the RRP community, where patients have long suffered from the physical and psychological toll of recurring airway surgeries. By shifting the paradigm from surgical removal to immunological control, Inovio is tapping into a significant unmet need.
However, the hardware requirements highlight a growing trend in regulatory oversight: as digital and electronic drug-delivery devices become more complex, the FDA is increasingly treating the "device" and the "drug" as an inseparable ecosystem. The focus on "human factors"—the study of how clinicians interact with technology—indicates that the FDA is prioritizing the prevention of "use error" as a central pillar of patient safety.
For Inovio, the next 18 months will be a period of dual-track execution: scaling up a complex commercial supply chain while simultaneously performing rigorous forensic engineering on the Cellectra 5PSP. If the company can navigate these requirements successfully, Voceryq could serve as a blueprint for future DNA-based therapies. If the studies reveal significant design flaws, however, the company may face the expensive and time-consuming prospect of a mid-launch hardware redesign.
As the industry watches, the success of Voceryq will be measured not just by its ability to keep patients out of the operating room, but by the reliability and safety of the device that brings it into their cells.
About the Author