In a milestone for genomic medicine, the U.S. Food and Drug Administration (FDA) has granted accelerated approval to Inovio Pharmaceuticals for Voceryq (doruxapapogene ralaplasmid and lalikinogene sifuplasmid-gxfd). This approval marks the first time a plasmid DNA-based immunotherapy has cleared the federal regulatory hurdles, offering a potential breakthrough for adults suffering from recurrent respiratory papillomatosis (RRP).
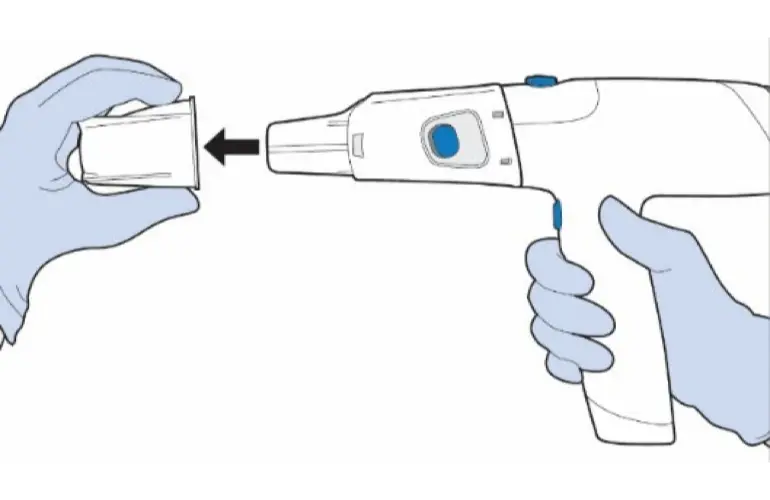
However, the approval comes with strings attached. The FDA has issued specific mandates requiring Inovio to conduct rigorous postmarketing studies concerning the Cellectra 5PSP, the specialized electroporation delivery device required to administer the therapy. While the clinical efficacy of the drug itself has satisfied regulators, the agency remains focused on the human-machine interface and safety profile of the delivery hardware.
The Intersection of Genetic Innovation and Device Usability
Voceryq represents a sophisticated approach to treating RRP, a rare, debilitating condition caused by human papillomavirus (HPV) types 6 and 11. These low-risk HPV strains trigger persistent, non-cancerous growths in the airway, often necessitating frequent, invasive surgeries to keep the patient’s breathing passages clear.
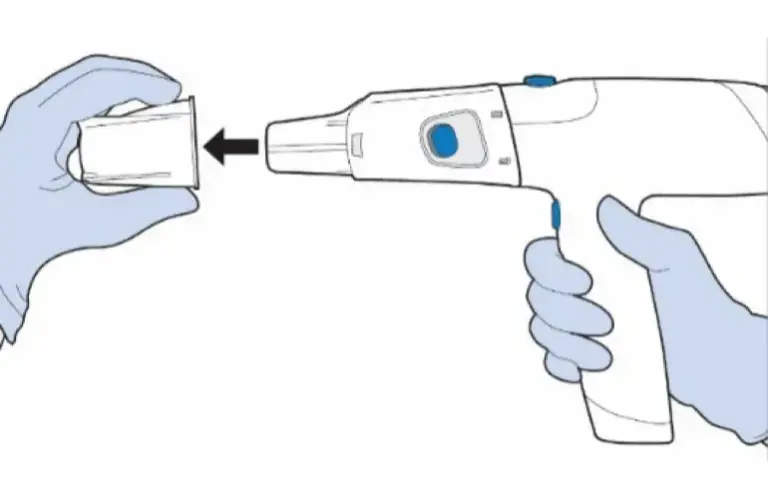
The treatment works by introducing two DNA plasmids—circular loops of genetic material—into the patient’s muscle cells. Once inside, the Cellectra 5PSP device delivers a controlled, 10-second burst of electrical pulses, a process known as electroporation. This brief current temporarily creates microscopic pores in the cell membranes, allowing the DNA to enter. Once inside, the plasmids provide the instructions necessary for the body to prime its T cells, effectively teaching the immune system to identify and neutralize the papilloma-infected cells.
Despite the therapeutic elegance of the drug, the FDA has identified "open questions" regarding the usability of the Cellectra 5PSP. Specifically, the regulator has demanded a comprehensive reanalysis of Inovio’s human factors validation studies. The agency’s 2026 guidance emphasizes the necessity of distinguishing between "close calls"—where a user realizes a mistake before harm occurs—and actual use errors. Inovio must now perform a deep-dive root cause analysis on these interactions to ensure that the device’s labeling, training protocols, and mechanical design do not pose a risk to healthcare providers or patients.
Furthermore, the FDA is requiring Inovio to verify and validate the effectiveness of the device’s "sharps injury prevention" features. Because the procedure involves an injection followed by an electrical delivery sequence, the risk of accidental needle sticks is a paramount concern for clinical staff.
Chronology: A Decade of Regulatory Navigation
The path to Voceryq’s approval is the culmination of years of iterative development and regulatory engagement.
- October 2016: Inovio encountered significant regulatory friction regarding its earlier DNA immunotherapy candidate, VGX-3100. The FDA placed a clinical hold on the Phase 3 program, specifically citing concerns over the shelf-life and sterility of the disposable components of the Cellectra delivery device.
- June 2017: After addressing the FDA’s requirements regarding device component integrity, Inovio successfully had the hold lifted, clearing the way for further trials. This period served as a formative learning experience for the company, underscoring the FDA’s insistence that the delivery hardware be held to the same standard of scrutiny as the drug payload itself.
- October 30, 2026: The FDA officially granted accelerated approval to Voceryq for the treatment of RRP in adults. On this same day, Inovio announced its commercial roadmap, projecting that orders would begin within 60 days, with a full-scale commercial launch slated for the first quarter of 2027.
- Post-Approval Mandates (Present): Inovio is now operating under a strict timeline to address the postmarketing commitments. The company must submit final reports for its human factors reanalysis and sharps injury prevention verification by June 30, 2027. If these studies indicate that the device requires design modifications, Inovio must submit a supplemental validation protocol, followed by a formal prior approval supplement by December 30, 2027.
The Evidence Base: Efficacy and Clinical Rationale
The accelerated approval for Voceryq was predicated on its ability to address an "intermediate clinical endpoint"—specifically, the frequency of surgical interventions required to maintain airway patency.
In a pivotal open-label, single-arm clinical trial involving 32 adults, researchers observed promising results. Among the 26 patients included in the FDA’s primary efficacy analysis—all of whom had a history of at least three surgeries in the 12 months preceding the trial—27% required zero surgical interventions in the year following treatment.
A subsequent retrospective follow-up study provided further encouragement, suggesting durability. Out of 28 participants, 14 required no surgery during the second year post-treatment. While these results are statistically significant in the context of a rare disease, the FDA has mandated a large-scale confirmatory study to verify these findings. This long-term study must be completed by August 30, 2032. The agency retains the authority to rescind approval if the trial fails to demonstrate durable clinical benefit or if the company fails to move with due diligence.
The treatment regimen itself is rigorous. After a surgeon clears the airway of visible growths, the patient receives a 1 mL dose every three weeks for four doses. Because of the electroporation process, patients must be monitored for at least 30 minutes following each session to ensure no adverse local reactions occur.
Official Responses and Strategic Implications
Inovio has framed the FDA’s requirements as a routine component of the regulatory lifecycle for novel delivery platforms. In an email correspondence, an Inovio spokesperson stated, "Postmarketing commitments are a normal part of the approval process, and we look forward to working with the agency to complete them. We do not expect them to affect our plans to make Voceryq available to order and ship in approximately 60 days."
From an industry perspective, this development highlights the evolving landscape of "combination products"—therapies that rely on both a drug and a medical device for efficacy. As the FDA updates its guidance on human factors and usability engineering, developers of such products must be prepared for increased oversight. The burden of proof is no longer just about whether the drug works, but whether the average clinician can safely and effectively navigate the complex interplay between the drug and the device in a busy hospital or clinic setting.
Future Outlook: Challenges and Opportunities
The successful deployment of Voceryq will hinge on two primary factors: the clinical adoption of the Cellectra 5PSP device and the successful completion of the required postmarketing studies.
For the medical community, the transition from repetitive surgical management to a DNA-based immunotherapy represents a shift in the standard of care for RRP. However, this transition requires training. Only healthcare providers specifically trained on the Cellectra 5PSP are authorized to administer Voceryq. Furthermore, because the device is ordered separately from the medication, the logistics of procurement will be a critical factor for hospital pharmacies and surgical centers.
Should Inovio successfully navigate the device-related hurdles by the 2027 deadlines, Voceryq could serve as a blueprint for the future of DNA immunotherapies. By demonstrating that the platform can effectively treat RRP while managing the risks associated with electroporation, Inovio may open the door for similar therapies targeting more common or complex conditions.
Conversely, any failure to address the FDA’s concerns regarding device usability or sharps safety could complicate the commercial rollout. The strict timeline set by the agency serves as a reminder that for cutting-edge medical technologies, the "product" is an inseparable combination of the medicine and the machine. As the industry moves toward 2027, the medical community will be watching closely to see if Inovio can solidify the safety of its delivery system while simultaneously proving the long-term, life-altering efficacy of its pioneering immunotherapy.
About the Author