For decades, the medical community has grappled with the devastating, inexorable progression of Huntington’s disease (HD). This fatal neurodegenerative disorder, characterized by a complex array of motor, cognitive, and psychiatric symptoms, has long presented a puzzle to researchers: why do different regions of the brain succumb to the disease at different rates, and what is the precise "molecular trigger" that turns an inherited genetic predisposition into cellular catastrophe?
In a significant leap forward for neurology, a collaborative team of scientists—led by researchers at the Broad Institute of MIT and Harvard, Harvard Medical School, and McLean Hospital—has published a landmark study in the journal Cell. This research builds upon a 2025 revelation that fundamentally altered the field’s understanding of HD, providing a potential roadmap for the next generation of therapeutic interventions.
The Core Discovery: A Molecular Threshold of Toxicity
The foundational premise of Huntington’s disease has historically rested on the HTT gene. Individuals who inherit a version of this gene featuring an expanded "CAG" repeat—a three-letter DNA sequence repeated 40 or more times—are at high risk for developing the disease. For years, the prevailing wisdom suggested that the mere presence of these repeats was the primary driver of pathology.
However, the 2025 research from the Broad Institute team upended this narrative. Their investigations revealed that in the striatum—the brain region most severely impacted by Huntington’s—the disease process is not a linear, constant decline. Instead, it is a cumulative, slow-burn phenomenon. The HTT mutation only reaches a toxic state once the DNA repeats expand significantly, accumulating 150 or more CAG units.
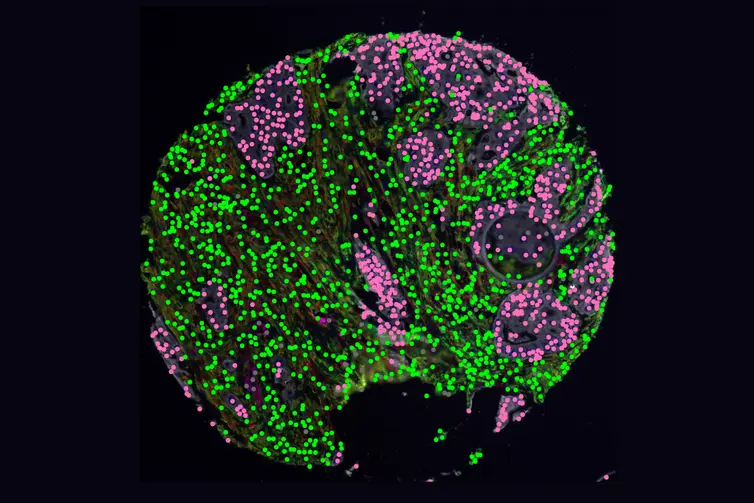
The new study, published in 2026, confirms that this "threshold of toxicity" is not a localized quirk of the striatum. By analyzing more than 125,000 cells from neocortex samples of eleven deceased Huntington’s patients, the research team discovered that the exact same mechanism drives pathology in the neocortex, a region critical for higher-order brain functions.
Chronology of the Research: From Mystery to Mechanism
The 2025 Turning Point
The 2025 study was the "aha!" moment for the field. By utilizing advanced single-cell sequencing, researchers were able to observe how HTT repeats behave over the course of a human lifespan. They identified that the DNA repeats are not static; they expand slowly throughout a person’s life. When these repeats cross the 150-mark threshold, the cell enters a state of molecular dysfunction, leading to the characteristic symptoms of Huntington’s.
The 2026 Expansion
While the 2025 findings were revolutionary, they left a critical question unanswered: Was this striatal expansion an anomaly, or did it explain the damage seen elsewhere in the brain? To answer this, the team turned to the neocortex. Using the same sophisticated high-resolution imaging and transcriptomic analysis, they mapped the repeat lengths in cortical cells.
The findings were striking. Just as in the striatum, cortical cells began to exhibit markers of severe damage only after the HTT gene surpassed the 150-repeat threshold. However, the researchers noted a vital difference in "kinetics": the neocortex cells reached this threshold much more slowly than those in the striatum. This difference in the speed of repeat expansion provides a compelling explanation for why some brain regions degrade faster than others, and why the clinical presentation of the disease can vary so significantly between patients.
Supporting Data: Understanding the Cellular Landscape
The researchers’ analysis of 125,000 cells provided a granular look at the disease process that had previously been invisible to science. Beyond the repeat threshold, the study uncovered two major pieces of evidence that reshape our understanding of the cellular environment in an HD brain:
1. Divergent Gene Expression
While the trigger (the 150-repeat threshold) is the same in both the striatum and the neocortex, the consequences are not. The team observed that the HTT mutation triggers different gene expression patterns in cortical cells compared to striatal cells. This suggests that while the root cause of the toxicity is a shared mechanism, the downstream impact on cellular machinery is context-dependent. This nuance may explain why HD patients experience such a diverse range of motor versus cognitive symptoms.
2. The Truth About Nuclear Inclusions
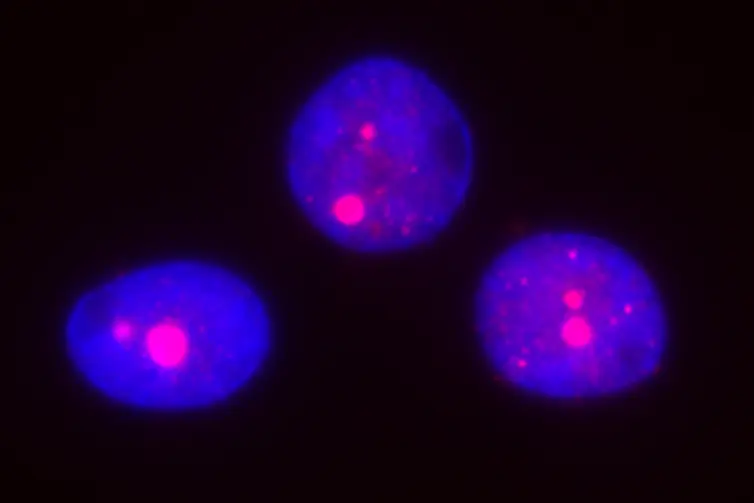
One of the most enduring debates in neurobiology involves "nuclear inclusions"—clumps of mutant Huntingtin protein that accumulate within the cell nucleus. For decades, scientists have debated whether these inclusions are the "executioners" of the cell, an irrelevant byproduct, or even a protective mechanism meant to sequester toxic proteins.
The 2026 study provides a clearer, albeit sobering, answer. Nuclear inclusions were found to be exceptionally rare and only present in cells that had far exceeded the 150-repeat threshold. These cells were also exhibiting extreme patterns of abnormal gene expression. Consequently, the authors conclude that these inclusions are not a defensive structure, but rather a "red flag"—a late-stage marker that the neuron is in a terminal state of dysfunction and is nearing cell death.
Official Responses and Expert Perspective
The implications of these findings have sent a ripple of optimism through the neurological community, particularly regarding the development of drugs that can "intercept" the repeat expansion process.
"These new results suggest that such medicines, if and when they’re successfully developed, might be able to address not just one of the pathologies in the disease, but the two big ones," noted Steve McCarroll, co-senior author of the study and an institute member at the Broad.
The urgency of these findings is emphasized by Sabina Berretta, co-senior author and director of the Harvard Brain Tissue Resource Center. "By pointing to a shared disease mechanism across brain regions, our findings suggest these new efforts are on the right track. We hope this evidence adds urgency to developing such treatments," Berretta stated.
The researchers were also quick to acknowledge the human cost of the research and the immense contribution of the patients involved. "Above all, this work is a testament to the extraordinary generosity, commitment, and foresight of brain donors and their families, whose contributions made these discoveries possible and to whom we are deeply grateful," Berretta added.
Implications for Future Medicine: A New Horizon
The realization that a single, universal mechanism—the expansion of HTT repeats beyond a specific threshold—is responsible for the damage across multiple brain regions is a game-changer for drug discovery.
Targeting the Mechanism
Current therapeutic efforts are increasingly focused on gene-editing technologies and antisense oligonucleotides (ASOs) designed to stabilize the HTT gene or lower the levels of the toxic protein. If these treatments can successfully slow the expansion of repeats before they hit the 150-threshold, they could theoretically prevent the neurodegeneration from ever initiating.
A Unified Therapeutic Strategy
Prior to this study, scientists feared they might need to develop region-specific therapies to treat the striatum versus the cortex. This latest research suggests a more streamlined approach: a systemic treatment that targets the repeat expansion mechanism could potentially protect the entire brain, addressing both the motor symptoms (associated with the striatum) and the cognitive decline (associated with the neocortex).
The Path Forward
The path from "mechanistic discovery" to "clinical application" remains long. Challenges regarding the delivery of genetic medicines across the blood-brain barrier and ensuring long-term safety remain. However, the discovery that there is a defined "point of no return" (the 150-repeat threshold) provides researchers with a clear biological target.
By identifying that these cells remain viable for a significant period before hitting this threshold, the research highlights a "window of opportunity" for intervention. For millions affected by Huntington’s disease and their families, this shift from understanding the disease as an inevitable decay to viewing it as a targetable, threshold-based process represents the most promising development in the field in over a generation.
As the scientific community shifts its focus toward these new targets, the focus will now turn to refining the delivery of therapies that can prevent this expansion. The work conducted by the teams at the Broad, Harvard, and McLean serves as a powerful reminder that when it comes to neurodegenerative diseases, the key to a cure lies in decoding the quiet, molecular clockwork that dictates the health of our neurons.
About the Author