In a significant stride for interventional cardiology, Teleflex Incorporated has announced that its Freesolve Resorbable Magnesium Scaffold (RMS) has received Breakthrough Device Designation from the US Food and Drug Administration (FDA). This regulatory milestone is a pivotal step toward bringing a "leave-nothing-behind" therapy to the US market, potentially transforming how clinicians manage complex coronary artery disease.
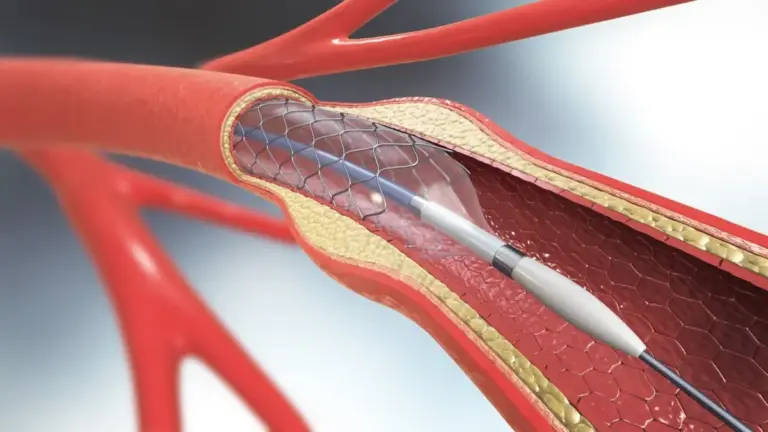
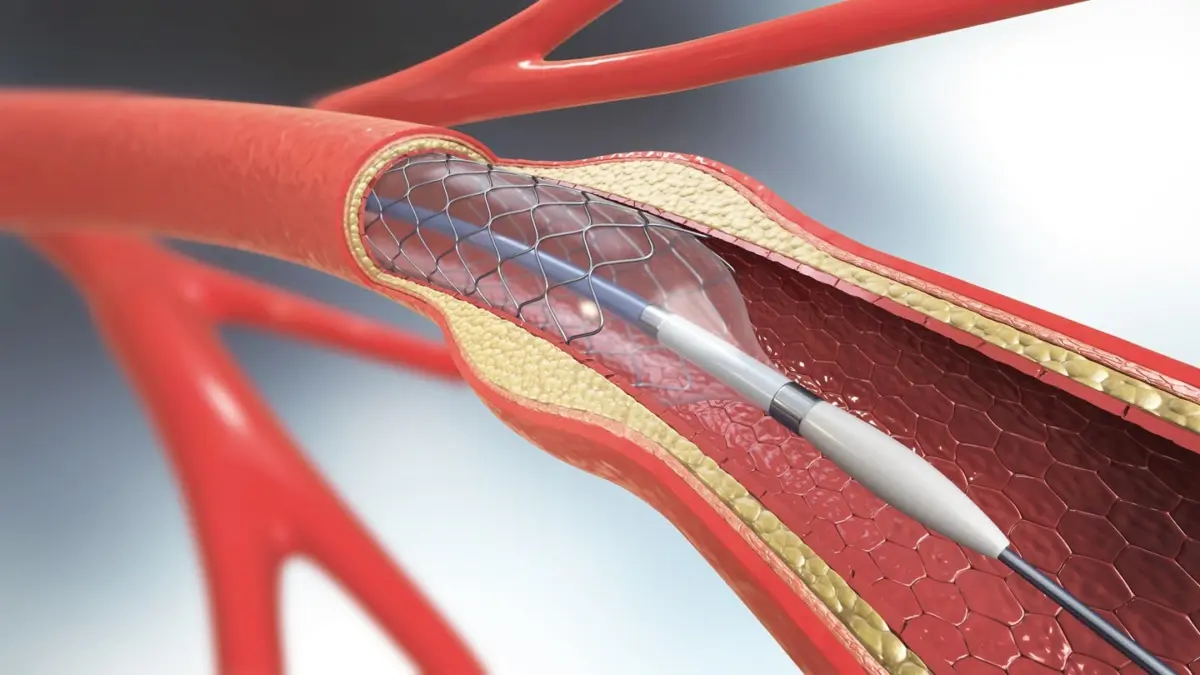
The Freesolve RMS is engineered to address the limitations of traditional permanent metallic stents. By utilizing advanced metallurgical and polymer science, Teleflex aims to provide temporary structural support to coronary arteries before being safely resorbed by the body.
Main Facts: Redefining Coronary Scaffolding
The FDA’s Breakthrough Device Designation is reserved for medical technologies that provide for more effective treatment or diagnosis of life-threatening or irreversibly debilitating conditions. By granting this status to the Freesolve RMS for percutaneous coronary intervention (PCI), the FDA acknowledges that this technology may offer significant advantages over current standards of care.
At its core, the Freesolve RMS integrates two proprietary technologies:
- BIOmag Magnesium Alloy: A high-strength, biocompatible material that serves as the scaffold’s structural backbone.
- BIOlute Bioabsorbable Coating: A sophisticated coating designed to facilitate controlled drug delivery and minimize inflammatory responses during the healing process.
The primary clinical advantage of the Freesolve system is its transient nature. Unlike conventional drug-eluting stents (DES) that remain in the artery for a lifetime, the Freesolve scaffold is designed to be resorbed by the body within 12 months. Current data suggests that approximately 99% of the magnesium alloy is cleared by the body within this timeframe. This "vanishing" act is intended to restore natural vessel vasomotion—the ability of the artery to expand and contract—thereby potentially reducing the long-term risk of late-stage complications such as neoatherosclerosis or stent thrombosis.
Chronology: A Trajectory of Clinical Advancement
The development of the Freesolve RMS is the culmination of years of rigorous research and iterative design improvements.
Early Development and CE Mark
Before targeting the US market, Teleflex focused on establishing clinical safety and efficacy in international territories. The device successfully secured the CE mark, allowing it to enter markets across Europe, the Middle East, Asia-Pacific, and South America. These international commercial experiences have served as a real-world evidence base, confirming the scaffold’s performance in diverse patient populations.
The BIOMAG-I Study
The clinical narrative began in earnest with the BIOMAG-I First-In-Human (FIH) study. This trial was critical in establishing the safety profile of the magnesium-based technology. The study followed patients over an extended period, yielding compelling long-term results. Most notably, researchers reported a target lesion failure (TLF) rate of just 3.5% at the four-year mark. Perhaps more impressively, the study recorded no new clinical events between the second and fourth years, suggesting that once the scaffold is resorbed and the vessel heals, the patient’s clinical outlook remains stable.
The Peripheral Pivot
In 2024, the FDA recognized the potential of the Freesolve technology for a different indication: chronic limb-threatening ischemia (CLTI). The agency granted Breakthrough Device Designation for the Freesolve below-the-knee (BTK) scaffold. This earlier designation provided Teleflex with a roadmap for navigating the US regulatory environment, creating a foundational understanding of the agency’s expectations for the magnesium-based platform.
The BIOMAG-III Pivotal Trial
Currently, Teleflex is conducting the BIOMAG-III Pivotal Trial. This large-scale, prospective study is the definitive evaluation required to support a Premarket Approval (PMA) application with the FDA. The trial is meticulously designed to provide the clinical evidence necessary to demonstrate that the Freesolve RMS is not only safe but also provides a clear clinical benefit compared to existing technologies.
Supporting Data: Why Magnesium Matters
The shift toward bioresorbable scaffolds (BRS) represents a fundamental change in philosophy in interventional cardiology. For decades, the "gold standard" has been the permanent metallic stent. While highly effective at keeping arteries open, these permanent implants are inherently "foreign bodies" that can remain a permanent site for chronic inflammation.
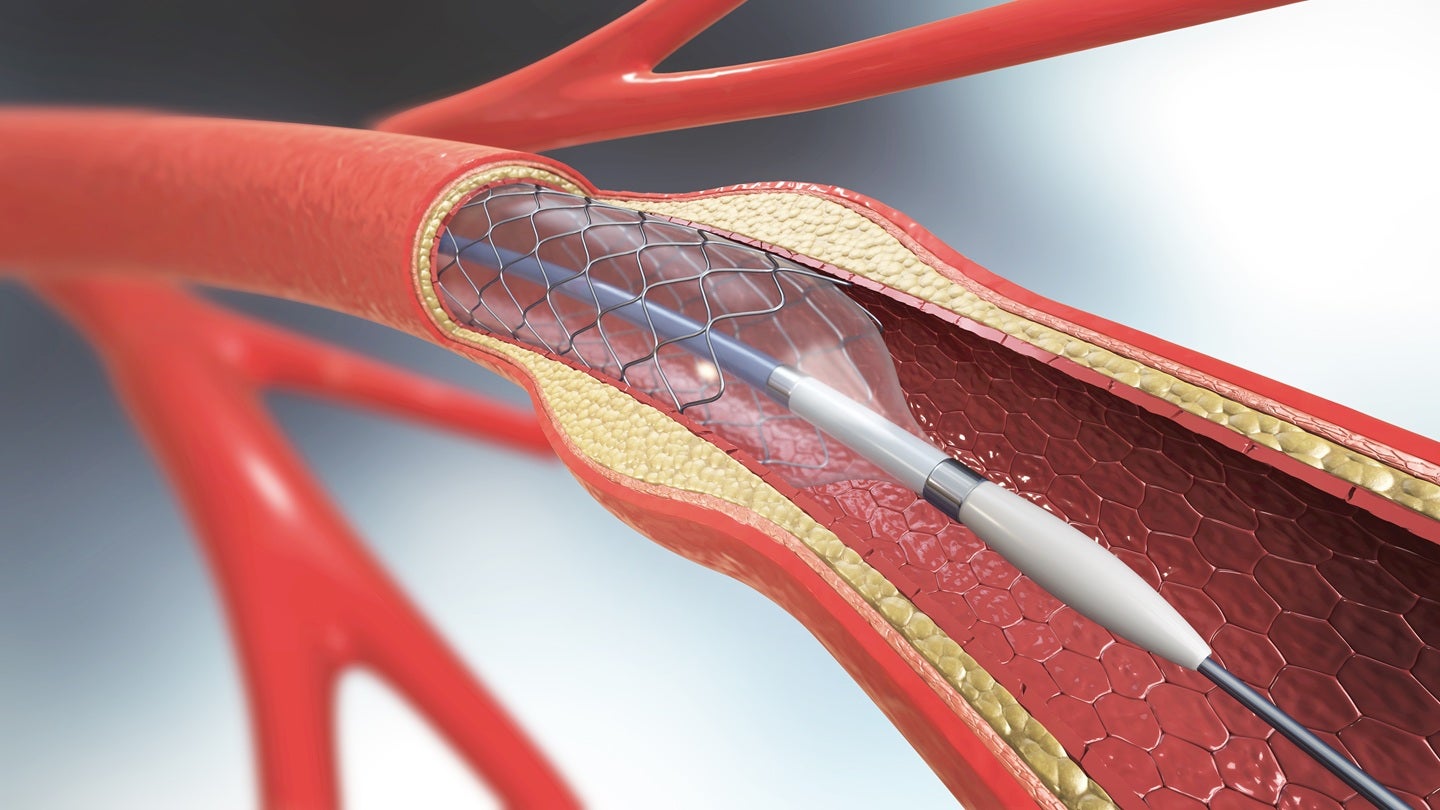
The clinical data from the BIOMAG-I trial provides a glimpse into why the medical community is optimistic about the Freesolve platform:
- Mechanical Integrity: The BIOmag alloy provides sufficient radial strength to support the vessel during the acute healing phase following PCI.
- Bio-Resorption Kinetics: The transition from a rigid scaffold to a fully resorbed state occurs in a predictable, controlled manner, minimizing the mechanical stress on the vessel wall during the degradation process.
- The "Leave-Nothing-Behind" Benefit: By eliminating the permanent metal cage, the Freesolve system aims to allow for the future physiological remodeling of the coronary artery. In theory, this allows patients to remain candidates for future interventions, including bypass surgery, should their disease progress—a significant advantage over patients who have a full length of their artery permanently "caged" by metallic stents.
Official Responses: Strategic Vision
The leadership at Teleflex has framed this designation as more than just a regulatory hurdle; they see it as a validation of their long-term strategic commitment to vascular innovation.
Jason Fontana, President and Interventional General Manager at Teleflex, emphasized the importance of this milestone: "At Teleflex, we are advancing vascular intervention through strategic investment and continuous innovation. This designation marks an important milestone in our efforts to bring Freesolve RMS to patients in the US."
Fontana further highlighted the cross-application potential of their platform, noting: "Building on the FDA’s previous breakthrough device designation for Freesolve RMS in below-the-knee applications, it further underscores the potential of bioresorbable magnesium scaffold technology and our commitment to a leave-nothing-behind approach. Freesolve RMS has the potential to redefine clinical practice in today’s cath labs."
The emphasis on "redefining clinical practice" suggests that Teleflex is not merely looking to offer an alternative to existing stents, but is aiming to shift the standard of care for a significant subset of patients suffering from coronary artery disease.
Implications: The Future of the Cath Lab
The implications of the Freesolve RMS breakthrough designation are far-reaching, touching upon clinical, economic, and procedural aspects of cardiology.
Clinical Implications
For patients, the prospect of a device that disappears is highly attractive. Many patients are apprehensive about the idea of living with permanent implants. If the BIOMAG-III trial validates the long-term safety seen in the BIOMAG-I study, cardiologists may have a potent tool to offer patients who are younger or have less complex lesion morphologies where a permanent stent might be considered an "over-treatment."
Economic and Regulatory Implications
The FDA’s Breakthrough Device Program is designed to accelerate the timeline for regulatory review. By granting this status, the FDA agrees to provide intensive guidance to the manufacturer on efficient development, which often includes more frequent interactions with agency staff. For Teleflex, this could shave months or even years off the time it takes to move from the end of the BIOMAG-III trial to actual commercial launch. This expedited timeline is crucial in the competitive landscape of medical device manufacturing, where the pace of innovation is relentless.
Changing the Paradigm of PCI
The "leave-nothing-behind" movement in cardiology has faced setbacks in the past, largely due to the complications associated with early-generation bioresorbable scaffolds (many of which were made of polymers). The failure of those early devices to match the performance of metallic stents cast a shadow over the entire BRS category. However, the use of magnesium alloy in the Freesolve device represents a technological "re-start." Magnesium is naturally occurring in the body, potentially offering a more favorable degradation profile than the polymers used in previous generations of BRS.
If the Freesolve RMS proves successful, it will likely lead to a resurgence in interest regarding resorbable technology. It would signal to the industry that the limitations of the past—namely, scaffold thickness and unpredictable degradation—have been solved through metallurgical innovation.
Conclusion
The FDA Breakthrough Device Designation for the Teleflex Freesolve RMS is a significant vote of confidence in the future of resorbable vascular technology. As the company moves through the BIOMAG-III Pivotal Trial, the eyes of the cardiology world will be fixed on the data. If the trial delivers on its promise, the Freesolve system could well become a cornerstone of modern interventional cardiology, offering a sophisticated, temporary solution for patients who deserve the most advanced care available. By blending the durability required for initial scaffolding with the grace of complete bioresorption, Teleflex is positioning itself at the vanguard of a movement that seeks to treat the disease while leaving the patient’s anatomy as close to its natural state as possible.
About the Author