The landscape of modern pharmacology shifted fundamentally in May 2026. With the U.S. FDA’s landmark approval of vepdegestrant (Veppanu®)—the first-ever Proteolysis Targeting Chimera (PROTAC) to reach the clinic—the pharmaceutical industry validated a modality that had long remained a theoretical promise. By successfully targeting and degrading the estrogen receptor (ER) in patients with ER-positive, HER2-negative, ESR1-mutated metastatic breast cancer, vepdegestrant proved that "chemical knockdown" is not merely a laboratory curiosity, but a viable clinical strategy.
However, the success of vepdegestrant belies a harsh reality: PROTACs are notoriously difficult to formulate as oral medicines. As drug developers race to apply this modality to a wider array of "undruggable" proteins, they are colliding with the stubborn physical laws governing pharmacology. Achieving the necessary oral exposure for these large, complex molecules requires a fundamental rethink of medicinal chemistry and preclinical strategy.
The Mechanism: Why PROTACs Defy Traditional Rules
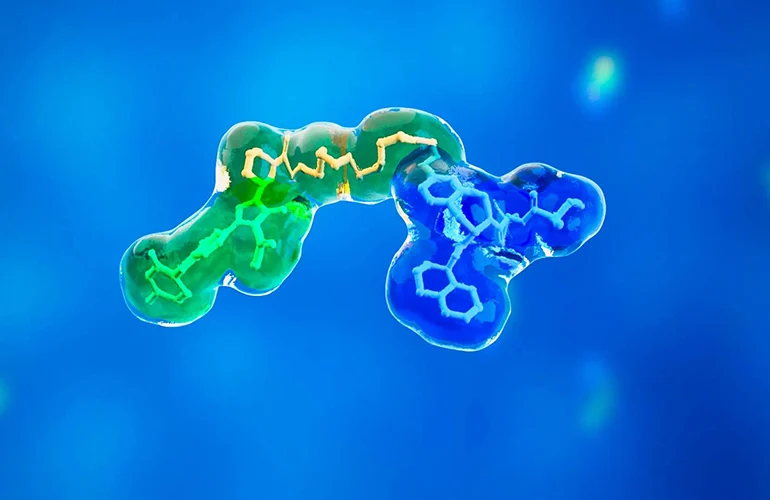
To understand the challenge, one must first understand the mechanism. PROTACs are heterobifunctional molecules consisting of three distinct domains: a warhead that binds the target protein, a ligand that recruits an E3 ubiquitin ligase, and a chemical linker connecting the two.
Unlike traditional inhibitors, which must occupy a protein’s active site in a 1:1 stoichiometric ratio, PROTACs operate catalytically. They induce the proximity-driven ubiquitination of the target, effectively marking it for destruction by the cell’s internal waste-disposal unit, the ubiquitin-proteasome system. This "event-driven" mechanism allows for sustained biological impact even at low drug concentrations, theoretically offering a way to target proteins that lack traditional binding pockets.
Yet, the structural complexity required for this "molecular handshake" creates a significant hurdle. Because they must bridge two proteins, PROTACs are inherently large, typically exceeding 700 Da, with high polar surface areas and numerous rotatable bonds. These characteristics place them squarely in the "beyond-Rule-of-Five" (bRo5) chemical space, where traditional guidelines for oral bioavailability—such as Lipinski’s Rule of Five—often fail to predict clinical success.
Chronology: From Concept to Clinical Reality
The journey to the first approved PROTAC was a decade-long endeavor that transformed early academic breakthroughs into a high-stakes commercial pipeline.
- 2010s: The Early Validation Phase. Researchers demonstrated that PROTACs could effectively degrade disease-relevant proteins in vitro. However, the initial focus was on refining the chemical architecture to ensure target specificity.
- 2020–2023: The Optimization Pivot. As the modality moved toward human testing, companies began to hit "the wall" of oral exposure. It became clear that potency in a cell culture did not translate to systemic circulation in vivo. This period saw a shift toward systematic studies of linker rigidity, E3 ligase selection, and metabolic stability.
- 2024: A New Framework. Industry analysis began to move away from binary pass/fail screening. Researchers identified that properties such as solvent-exposed hydrogen-bond donors (eHBD) and conformational folding were more predictive of oral success than molecular weight alone.
- 2026: The Regulatory Threshold. The approval of vepdegestrant served as the industry’s "North Star," confirming that with sufficient optimization of the linker and ligase-recruitment strategies, orally bioavailable degraders could meet stringent FDA safety and efficacy standards.
Supporting Data: The Anatomy of the Exposure Problem
The struggle for oral bioavailability is essentially a struggle against three interconnected physical liabilities: poor aqueous solubility, limited membrane permeability, and rapid metabolic clearance.
The eHBD Metric
Traditional metrics assume that high molecular weight equals poor absorption. However, recent data suggests that the number of solvent-exposed hydrogen-bond donors (eHBD) is a far more critical discriminator. Studies have indicated that keeping eHBD ≤ 2 is a key threshold for maintaining the membrane permeability required for oral administration.
The Role of Biorelevant Media
A common pitfall is relying on simple aqueous buffers for solubility testing. PROTACs often behave differently in the complex environment of the gastrointestinal tract. Research has shown that many degraders exhibit improved solubility in fasted- and fed-state simulated intestinal fluids (FaSSIF and FeSSIF). Consequently, the "food effect"—the observation that a drug’s absorption is altered by the presence of food—is not just an outcome to be managed, but a variable to be engineered early in the program.
Linker Dynamics
The linker is the most versatile, yet most dangerous, part of the molecule. Replacing standard PEG (polyethylene glycol) linkers with more rigid structures, such as 1,4-disubstituted phenyl rings, has been shown to reduce excessive rotatable bonds and improve passive membrane permeability. Furthermore, modifying the linker can hide polar groups within a "folded" conformation, effectively shielding the molecule from the aqueous environment during transport across the gut wall.
Addressing the Challenges: Strategies for Success
To move beyond the limitations of current PROTAC designs, experts advocate for a multi-disciplinary approach that integrates DMPK (Drug Metabolism and Pharmacokinetics) profiling into the earliest stages of hit-to-lead optimization.
1. Strategic E3 Ligase Selection
Not all E3 ligases are created equal regarding developability. While Cereblon (CRBN) and Von Hippel-Lindau (VHL) are the industry standards, CRBN-based degraders often allow for more compact molecular architectures. As seen with vepdegestrant, selecting the right ligase is as much a decision about the molecule’s physical properties as it is about the target biology.
2. Engineering Intramolecular Hydrogen Bonds
One of the most sophisticated approaches to improving permeability is designing molecules that "self-shield." By carefully placing hydrogen-bond donors and acceptors, chemists can encourage the PROTAC to adopt a folded, low-energy conformation. This reduces the exposed polar surface area, allowing the molecule to navigate the lipophilic environment of the cell membrane more efficiently.
3. The Prodrug Path
When structural modifications reach their limit, the prodrug approach offers an escape hatch. By attaching a lipophilic "mask" to the active PROTAC, developers can improve solubility and absorption, with the mask being cleaved by systemic enzymes once the molecule enters the bloodstream. While this adds complexity to the regulatory and manufacturing process, it may be the only way to make certain high-potency PROTACs viable for oral delivery.
4. Considering Molecular Glues
If a PROTAC remains too large or too polar, it is time to pivot. Molecular glues represent an adjacent modality. Unlike PROTACs, which use a linker, glues are smaller, more compact molecules that stabilize protein-protein interactions directly. While they require a different discovery strategy, they often start from a more favorable position in terms of drug-like properties.
Implications for Future Drug Discovery
The approval of vepdegestrant has sent a clear message to the pharmaceutical industry: the oral bioavailability barrier is not a wall, but a design challenge. The implication for future drug development is a necessary shift in culture.
Drug discovery teams can no longer afford to work in silos. Medicinal chemists, structural biologists, and DMPK experts must be aligned from the moment a target is identified. The goal is to build "developability" into the molecule from the outset. This includes:
- Fit-for-purpose assays: Moving away from generic cell-based assays toward those that specifically model intestinal absorption and metabolic stability.
- Early DMPK integration: Utilizing ultrasensitive bioanalysis to track exactly where and why exposure is lost, rather than waiting for late-stage preclinical failure.
- Translational awareness: Recognizing that the ultimate goal is not just a high-potency degrader, but a robust, shelf-stable oral medicine that can be dosed safely in humans.
Conclusion
The success of PROTACs in the clinical setting represents one of the most significant advancements in drug discovery in the 21st century. By hijacking the body’s natural protein disposal system, we have unlocked the ability to treat diseases that were previously thought to be beyond the reach of traditional medicine.
However, the "oral exposure problem" remains the defining bottleneck of the field. As we move forward, the winners in this space will be the companies that treat oral bioavailability as a core design parameter—leveraging structural biology, metabolic engineering, and creative chemistry to ensure that these powerful molecules can reach their targets effectively. The path forward is difficult, but for the millions of patients awaiting new therapeutic options, the reward for solving this puzzle is immeasurable.
Tao Xiong, Director of DMPK at WuXi AppTec, emphasizes that success in this field requires a holistic view of the molecule. With nearly two decades of experience in supporting over 200 global IND programs, Xiong maintains that the integration of permeability assessment platforms and early ADME profiling is the only way to successfully navigate the complex landscape of targeted protein degradation.
About the Author